Showing posts sorted by date for query RNA. Sort by relevance Show all posts

Wednesday, September 16, 2026

Glutamic Acid Helps Fresh-Cut Potatoes Stay Fresh by Silencing Browning Genes | Newswise

In continuation of my update on Glutamic acid 

The study identified a transcriptional regulatory module, StbZIP53-like2–StERF091, that suppresses the activity of key polyphenol oxidase genes linked to browning. By showing how glutamic acid activates this gene-control system, the work provides a clearer molecular basis for safer anti-browning strategies and may support the future development of quality-preserving treatments for fresh-cut potato products and other minimally processed vegetables.


Fresh-cut potatoes are increasingly popular because they fit modern food habits, but their shelf life is limited by rapid enzymatic browning after cutting. Existing methods to control browning include chemical, physical, and biological approaches, yet each has drawbacks. Some chemicals raise food-safety concerns, while physical treatments may cause off-flavors, and many biological methods are still difficult to commercialize. Earlier work had already shown that glutamic acid could reduce browning in fresh-cut potatoes, but the molecular mechanism remained unclear. That unresolved question made it necessary to investigate how glutamic acid regulates browning-related genes and upstream transcription factors in fresh-cut potato tissue.

A study (DOI:10.48130/ph-0026-0004) published in Plant Hormones on 20 March 2026 by Jingying Shi’s & Zunyang Song’s team, Shandong Agricultural University, demonstrates that glutamic acid-induced StbZIP53-like2 and StERF091 form a cooperative regulatory module that represses StPPO2 and StPPO3, thereby alleviating browning in fresh-cut potatoes.

The team first prepared fresh-cut potato shreds and compared untreated samples with samples immersed in a glutamic acid solution, then stored them at 4 °C and collected tissues across multiple time points. They measured ethylene production, performed RNA-seq and RT-qPCR analyses, and screened for transcription factors whose expression changed under glutamic acid treatment. Among 11 bZIP genes detected, StbZIP53-like2 stood out because its expression was strongly induced by glutamic acid. Subcellular localization assays showed that this protein is localized in the nucleus, consistent with its proposed role as a transcription factor. The researchers then examined whether StbZIP53-like2 directly controlled known browning-associated genes. Dual-luciferase assays showed that it repressed the promoter activity of StPPO3, while EMSA and yeast one-hybrid experiments confirmed direct binding to the C-box motif in the StPPO3 promoter. To identify interacting partners, the team screened a cDNA library and found StERF091, another glutamic acid-induced transcription factor. Yeast two-hybrid, GST pull-down, and co-immunoprecipitation assays together demonstrated that StbZIP53-like2 physically interacts with StERF091 both in vitro and in vivo. Like StbZIP53-like2, StERF091 was also localized in the nucleus. Next, the group tested the function of StERF091 in regulating browning genes. They found that StERF091 repressed the promoter activities of StPPO2 and StPPO3, but not StPPO7. EMSA and yeast one-hybrid assays further showed that StERF091 directly binds the GCC-box motifs in the StPPO2 and StPPO3 promoters. Most importantly, co-expression assays revealed that when StbZIP53-like2 and StERF091 acted together, repression of StPPO2 and StPPO3 became stronger than with either factor alone. This demonstrated that the two proteins form a functional inhibitory module that amplifies the anti-browning response triggered by glutamic acid.

Overall, the study shows that glutamic acid alleviates the browning of fresh-cut potatoes by inducing two transcriptional repressors, StbZIP53-like2 and StERF091, which cooperate to suppress StPPO2 and StPPO3. The findings move beyond a simple observation that glutamic acid works and explain why it works at the molecular level. This insight may help guide future preservation technologies aimed at extending the shelf life, visual quality, and commercial value of fresh-cut produce.


https://en.wikipedia.org/wiki/Glutamic_acid


Monday, August 31, 2026

FDA Approves Idvynso (doravirine and islatravir) for the Treatment of HIV-1 Infection in Adults

In continuation of my update on Doravirine

Merck (NYSE: MRK), known as MSD outside of the United States and Canada, announced the U.S. Food and Drug Administration (FDA) approval of  Idvynso, a new, two-drug single-tablet regimen of 100 mg doravirine and 0.25 mg islatravir, for the treatment of HIV-1 infection in adults to replace the current antiretroviral regimen in those who are virologically suppressed (HIV-1 RNA less than 50 copies per mL) on a stable antiretroviral regimen with no history of virologic treatment failure and no known substitutions associated with resistance to doravirine. Idvynso is contraindicated when co-administered with drugs that are strong cytochrome P450 (CYP)3A enzyme inducers and lamivudine (3TC) or emtricitabine (FTC). Co-administration with these drugs may decrease the effectiveness of Idvynso. See additional selected safety information on the following pages. Idvynso (pronounced ihd-VIHN-soh) will be available in pharmacies after May 11.

                                                                Islatravir


Doravirine


  • Idvynso is approved for adults with virologically suppressed HIV-1 with no history of virologic treatment failure and no known substitutions associated with resistance to doravirine
  • Idvynso is the first and only non-INSTI, tenofovir-free, once-daily, complete two-drug regimen to demonstrate non-inferior efficacy in a head-to-head Phase 3 trial versus three-drug regimen Biktarvyi (BIC/FTC/TAF)

“Advances in HIV treatment mean more people living with HIV are living longer — a remarkable achievement,” said Carl Baloney, Jr., president and chief executive officer of AIDS United. “People aging with HIV face additional health challenges, including managing multiple chronic conditions and medications at the same time. It is essential that management of HIV considers these factors in addition to virologic suppression when choosing an HIV treatment regimen.”

“Idvynso combines islatravir, a next-generation NRTI with multiple mechanisms of action, including translocation inhibition, with doravirine, an NNRTI with an established efficacy and safety profile. As the only two-drug, non-INSTI, tenofovir-free regimen, Idvynso expands therapeutic diversity beyond the currently available oral treatment options,” said Dr. Eliav Barr, senior vice president and chief medical officer, Merck Research Laboratories. “As the health needs of adults living with HIV change over time, Idvynso gives clinicians a new choice for HIV treatment. This approval marks an important new chapter in Merck’s long-standing commitment to research and discovery for people living with HIV.”

Idvynso is a complete regimen; co-administration with other antiretroviral medications for treatment of HIV-1 infection is not recommended. Severe skin reactions, including Stevens-Johnson syndrome/toxic epidermal necrolysis, have been reported with doravirine-containing regimens. Drug Rash with Eosinophilia and Systemic Symptoms was reported with Idvynso. Concomitant use of Idvynso and certain other drugs may result in known or potentially significant drug interactions, some of which may lead to loss of therapeutic effect of Idvynso and possible development of resistance, or possible clinically significant adverse reactions from greater exposures of a component of Idvynso. See additional selected safety information on the following pages.

“Idvynso is the first non-INSTI, tenofovir-free, two-drug regimen to demonstrate non-inferior efficacy to standard oral antiretroviral regimens, including Biktarvy. This makes Idvynso a potential alternative for people with virologically suppressed HIV who may need to switch their treatment,” said Dr. Amy Colson, director of research at Community Resource Initiative, Boston, Massachusetts.

Phase 3 studies supporting approval of Idvynso

The efficacy and safety of Idvynso is supported by Week 48 data from two randomized, active-controlled, non-inferiority trials [Trial 052 (NCT05630755) and Trial 051 (NCT05631093)] in virologically-suppressed (HIV-1 RNA less than 50 copies per mL) adults living with HIV. Participants must have been stably suppressed on their baseline regimen for at least 3 months prior to trial entry and had no history of treatment failure. Across the two trials, a total of 708 participants received once-daily Idvynso; of these, 81 (11%) participants were aged 65 years and older, including 10 (1%) aged 75 years and older.

In the double-blind Trial 052, participants were switched from Biktarvy [bictegravir/emtricitabine/tenofovir alafenamide (BIC/FTC/TAF)] to Idvynso. A total of 513 participants were randomized (2:1) and were switched to once-daily Idvynso (n=342) or remained on BIC/FTC/TAF (n=171). At baseline, participants had a mean age of 48 years (range: 19 to 77), 21% of participants were female, 61% were White, 31% were Black/African American, and 6% were Asian. A total of 23% identified as Hispanic/Latino.

In the open-label Trial 051, participants were switched from an oral ART (antiretroviral therapy) regimen to Idvynso. A total of 551 participants were randomized (2:1) and were switched to once-daily Idvynso (n=366) or remained on their baseline ART (bART) (n=185). Randomization was stratified by bART. At baseline, participants had a mean age of 50 years (range: 18 to 83), 40% of participants were female, 39% were White, 45% were Black/African American, and 5% were Asian. A total of 15% identified as Hispanic/Latino. At enrollment, 64% of the participants were receiving integrase strand transfer inhibitor (INSTI)-based regimens, 5% protease inhibitor (PI)-based regimens (including combinations with INSTI), and 30% were receiving other regimens.

https://en.wikipedia.org/wiki/Islatravir
https://en.wikipedia.org/wiki/Doravirine

Monday, January 5, 2026

FDA Approves Dawnzera (donidalorsen) as the First and Only RNA-targeted Prophylactic Treatment for Hereditary Angioedema

Ionis Pharmaceuticals, Inc. (Nasdaq: IONS) announced the U.S. Food and Drug Administration (FDA) approval of Dawnzera (donidalorsen) for prophylaxis to prevent attacks of hereditary angioedema (HAE) in adult and pediatric patients 12 years of age and older. Dawnzera is the first and only RNA-targeted medicine approved for HAE, designed to target plasma prekallikrein (PKK), a key protein that activates inflammatory mediators associated with acute attacks of HAE. Dawnzera 80mg is self-administered via subcutaneous autoinjector once every four (Q4W) or eight weeks (Q8W).


HAE is a rare and potentially life-threatening genetic condition that involves recurrent attacks of severe swelling (angioedema) in various parts of the body, including the hands, feet, genitals, stomach, face and/or throat. HAE is estimated to affect approximately 7,000 people in the U.S.

“Dawnzera represents a significant advance for people living with HAE who need improved treatment options. With strong and durable efficacy, convenient administration and the longest dosing option available, we believe Dawnzera will be the prophylactic treatment of choice for many people living with HAE. Importantly, the recently published switch data empowers patients and physicians with a roadmap for switching to Dawnzera from other prophylactic therapies,” said Brett P. Monia, Ph.D., chief executive officer, Ionis. “At Ionis, we are dedicated to turning groundbreaking science into life-changing medicines. With the early success of our first independent launch of TRYNGOLZA® for familial chylomicronemia syndrome (FCS), and now with Dawnzera, our second independent medicine approved in less than nine months, we are proudly delivering on that vision. To the patients, families, advocacy partners and investigators who helped make this moment a reality, we express our deepest gratitude.”

The approval of Dawnzera was based on positive results from the Phase 3 global, multicenter, randomized, double-blind, placebo-controlled OASIS-HAE study in patients with HAE. The study met its primary endpoint, with Dawnzera Q4W significantly reducing monthly HAE attack rate by 81% compared to placebo over 24 weeks. Mean attack rate reduction increased to 87% when measured from the second dose, a key secondary endpoint. Additionally, Dawnzera Q4W reduced moderate-to-severe HAE attacks by ~90% over 24 weeks when measured from the second dose.

These results are bolstered by the ongoing OASISplus open-label extension (OLE) study, in which Dawnzera Q8W had a similar effect as Q4W over time. Dawnzera demonstrated 94% total mean attack rate reduction from baseline across both dosing groups after one year in the OLE.

The OASISplus study also includes a switch cohort evaluating Dawnzera Q4W in patients previously treated with lanadelumab, C1-esterase inhibitor or berotralstat for at least 12 weeks. Switching to Dawnzera reduced mean HAE attack rate by 62% from prior prophylactic treatment over 16 weeks, with no mean increase in breakthrough attacks observed during the switch. A total of 84% of patients surveyed preferred Dawnzera over their prior prophylactic treatment, citing better disease control, less time to administer and less injection site pain or reactions.

Across clinical studies, Dawnzera demonstrated a favorable safety and tolerability profile. The most common adverse reactions (incidence ≥ 5%) were injection site reactions, upper respiratory tract infection, urinary tract infection and abdominal discomfort.

“As the first FDA-approved RNA-targeted therapy for HAE, Dawnzera represents a welcome advance in therapeutic options for preventing attacks. Today’s approval gives people living with HAE and their physicians another important choice for aligning treatment with individual needs,” said Anthony J. Castaldo, CEO & chairman of the board, U.S. Hereditary Angioedema Association (HAEA) and Hereditary Angioedema International (HAEi).

"People living with HAE manage this condition for all their lives, and many continue to face unpredictable, painful and dangerous breakthrough attacks even with current treatments. Durable efficacy is essential in maintaining long-term disease control,” said Marc Riedl, M.D., M.S., clinical director, U.S. HAEA Angioedema Center; University of California, San Diego; OASIS-HAE and OASISplus trial investigator. "Dawnzera is positioned to help meet patient needs, providing substantial and sustained reduction of HAE attacks, continued improvement over time and reduced burden of treatment.”

Dawnzera will be available in the U.S. in the coming days.

Ionis is committed to helping people access the medicines they are prescribed and will offer a suite of services designed to meet the unique needs of the HAE community through Ionis Every Step™. As part of Ionis Every Step, patients and healthcare providers will have access to a wide range of support and resources including dedicated support from a Patient Education Manager, assistance with the insurance approval process, information on affordability programs, access to the Dawnzera Direct digital companion and other ongoing services and resources to help patients stay on track. Visit Dawnzera.com for more information.

https://en.wikipedia.org/wiki/Donidalorsen

Thursday, July 21, 2022

FDA Approves Amvuttra (vutrisiran) for the Treatment of the Polyneuropathy of Hereditary Transthyretin-Mediated Amyloidosis in Adults


Alnylam Pharmaceuticals, Inc. (Nasdaq: ALNY), the leading RNAi therapeutics company, today announced that the U.S. Food and Drug Administration (FDA) approved Amvuttra (vutrisiran), an RNAi therapeutic administered via subcutaneous injection once every three months (quarterly) for the treatment of the polyneuropathy of hereditary transthyretin-mediated (hATTR) amyloidosis in adults. hATTR amyloidosis is a rare, inherited, rapidly progressive, and fatal disease with debilitating polyneuropathy manifestations, for which there are few treatment options. The FDA approval is based on positive 9-month results from the HELIOS-A Phase 3 study, where Amvuttra significantly improved the signs and symptoms of polyneuropathy, with more than 50 percent of patients experiencing halting or reversal of their disease manifestations.
“Twenty years ago, Alnylam was founded with the bold vision for RNA interference to make a meaningful impact on the lives of people around the world in need of new approaches to address serious diseases with significant unmet medical needs, such as hATTR amyloidosis. Today, Amvuttra has the potential to change the standard of care for people living with the polyneuropathy of this devastating disease,” said Yvonne Greenstreet, MBChB, Chief Executive Officer of Alnylam Pharmaceuticals. “We are so thankful to the patients, families and investigators involved in making Amvuttra a reality for the hATTR amyloidosis community. As the fifth RNAi therapeutic developed by Alnylam to receive regulatory approval in less than four years, we believe Amvuttra represents an important milestone that brings us one step closer to achieving our P5x25 goals aimed at Alnylam’s transition to a leading biotech company.”

The FDA approval of Amvuttra is based on positive 9-month results from HELIOS-A, a global, randomized, open-label, multicenter, Phase 3 study that evaluated the efficacy and safety of Amvuttra across a diverse group of patients with hATTR amyloidosis with polyneuropathy. 164 patients with hATTR amyloidosis were randomized 3:1 to receive either 25 mg of vutrisiran (N=122) via subcutaneous injection once every three months or 0.3 mg/kg of patisiran (N=42) via intravenous infusion once every three weeks (reference group) for 18 months. The efficacy of Amvuttra was assessed by comparing the Amvuttra group in HELIOS-A with the placebo group (n=77) from the landmark APOLLO Phase 3 study of patisiran, a randomized controlled study in a comparable patient population.
Amvuttra met the primary endpoint of the study, the change from baseline in the modified Neuropathy Impairment Score + 7 (mNIS+7) at 9 months. Treatment with Amvuttra (N=114) resulted in a 2.2 point mean decrease (improvement) in mNIS+7 from baseline as compared to a 14.8 point mean increase (worsening) reported for the external placebo group (N=67), resulting in a 17.0 point mean difference relative to placebo (p<0.0001); by 9 months, 50 percent of patients treated with Amvuttra experienced improvement in neuropathy impairment relative to baseline.

Amvuttra also met all secondary endpoints in the study at 9 months, with significant improvement in the Norfolk Quality of Life Questionnaire-Diabetic Neuropathy (Norfolk QoL-DN) score and timed 10-meter walk test (10-MWT), and improvements were observed in exploratory endpoints, including change from baseline in modified body mass index (mBMI), all relative to external placebo. Efficacy results at 18 months were consistent with 9-month data, with Amvuttra achieving statistically significant improvements compared to external placebo for all secondary endpoints including mNIS+7, Norfolk QoL-DN, 10-MWT and mBMI, and non-inferiority in serum TTR reduction relative to the within-study patisiran reference group.

-----------------------------------------------------------------------------------------------------------------


Friday, March 11, 2022

FDA Approves Leqvio (inclisiran), First-in-Class siRNA to Reduce Low-Density Lipoprotein Cholesterol (LDL-C)

                       In continuation of my update on inclisiran

                             Novartis today announced the US Food and Drug Administration (FDA) approval of Leqvio® (inclisiran), the first and only small interfering RNA (siRNA) therapy to lower low-density lipoprotein cholesterol (also known as bad cholesterol or LDL-C) with two doses a year, after an initial dose and one at three months.

"Leqvio is a revolutionary approach to lower LDL-C, and creates new possibilities for how  healthcare systems can impact cardiovascular disease, a defining public health challenge of our time," said Vas Narasimhan, Novartis CEO. "We now have the opportunity, working together with partners, to provide this first-ever approved LDL-C–lowering siRNA-based therapy to tackle ASCVD at scale across the United States."

Leqvio is indicated in the United States as an adjunct to diet and maximally tolerated statin therapy for the treatment of adults with clinical atherosclerotic cardiovascular disease (ASCVD) or heterozygous familial hypercholesterolemia (HeFH) who require additional lowering of LDL-C. The effect of Leqvio on cardiovascular morbidity and mortality is being explored in clinical trials currently underway.

"ASCVD is a substantial public health burden affecting 30 million Americans," said Norman Lepor, MD, a Los Angeles based cardiologist and a clinical investigator in the Phase III clinical program for Leqvio. "As a first-of-its-kind siRNA therapy, Leqvio works differently than other cholesterol treatments, with twice-yearly dosing that makes it a compelling option for the millions of people with ASCVD already on cholesterol-lowering medications struggling to reach their LDL-C target."

Leqvio reduces the amount of LDL-C in the bloodstream by improving the liver's natural ability to prevent the production of a protein that plays a role in keeping circulating cholesterol levels high6,7. It is a subcutaneous injection given by a healthcare provider with an initial dose, then again at three months, and then every six months1. This approach may help those who have trouble sticking to medicines that are self-administered and have greater dosing frequency. Leqvio will be available in early January 2022.

"People with ASCVD have most likely experienced a heart attack or stroke from high cholesterol, causing a burden on the family and having a negative impact on lives," said Andrea Baer, Executive Director of The Mended Hearts, Inc. "One of the first steps to improving patients' health is to manage high cholesterol and we're encouraged that this new twice-a-year treatment offers a new option." 

The FDA approval was based on results from the comprehensive Phase III ORION-9, -10 and -11 clinical trials, in which all 3,457 participants with ASCVD or HeFH had elevated LDL-C while receiving a maximally tolerated dose of statin therapy2,3. In the Phase III trials at month 17, Leqvio delivered effective and sustained LDL-C reduction of up to 52% vs. placebo and was reported to be well-tolerated with a safety profile shown to be comparable to placebo2,3. The most common side effects were mild to moderate injection site reaction (including pain, redness and rash), joint pain, urinary tract infection, diarrhea, chest cold, pain in legs or arms and shortness of breath2,3.

Novartis has obtained global rights to develop, manufacture and commercialize Leqvio under a license and collaboration agreement with Alnylam Pharmaceuticals, a leader in RNAi therapeutics.

https://en.wikipedia.org/wiki/Inclisiran

Wednesday, January 26, 2022

FDA Approves Leqvio (inclisiran), First-in-Class siRNA to Reduce Low-Density Lipoprotein Cholesterol (LDL-C)


Novartis  announced the US Food and Drug Administration (FDA) approval of Leqvio® (inclisiran), the first and only small interfering RNA (siRNA) therapy to lower low-density lipoprotein cholesterol (also known as bad cholesterol or LDL-C) with two doses a year, after an initial dose and one at three months.

"Leqvio is a revolutionary approach to lower LDL-C, and creates new possibilities for how healthcare systems can impact cardiovascular disease, a defining public health challenge of our time," said Vas Narasimhan, Novartis CEO. "We now have the opportunity, working together with partners, to provide this first-ever approved LDL-C–lowering siRNA-based therapy to tackle ASCVD at scale across the United States."

Leqvio is indicated in the United States as an adjunct to diet and maximally tolerated statin therapy for the treatment of adults with clinical atherosclerotic cardiovascular disease (ASCVD) or heterozygous familial hypercholesterolemia (HeFH) who require additional lowering of LDL-C. The effect of Leqvio on cardiovascular morbidity and mortality is being explored in clinical trials currently underway.

"ASCVD is a substantial public health burden affecting 30 million Americans," said Norman Lepor, MD, a Los Angeles based cardiologist and a clinical investigator in the Phase III clinical program for Leqvio. "As a first-of-its-kind siRNA therapy, Leqvio works differently than other cholesterol treatments, with twice-yearly dosing that makes it a compelling option for the millions of people with ASCVD already on cholesterol-lowering medications struggling to reach their LDL-C target."

Leqvio reduces the amount of LDL-C in the bloodstream by improving the liver's natural ability to prevent the production of a protein that plays a role in keeping circulating cholesterol levels high6,7. It is a subcutaneous injection given by a healthcare provider with an initial dose, then again at three months, and then every six months1. This approach may help those who have trouble sticking to medicines that are self-administered and have greater dosing frequency. Leqvio will be available in early January 2022.

"People with ASCVD have most likely experienced a heart attack or stroke from high cholesterol, causing a burden on the family and having a negative impact on lives," said Andrea Baer, Executive Director of The Mended Hearts, Inc. "One of the first steps to improving patients' health is to manage high cholesterol and we're encouraged that this new twice-a-year treatment offers a new option." 

The FDA approval was based on results from the comprehensive Phase III ORION-9, -10 and -11 clinical trials, in which all 3,457 participants with ASCVD or HeFH had elevated LDL-C while receiving a maximally tolerated dose of statin therapy2,3. In the Phase III trials at month 17, Leqvio delivered effective and sustained LDL-C reduction of up to 52% vs. placebo and was reported to be well-tolerated with a safety profile shown to be comparable to placebo2,3. The most common side effects were mild to moderate injection site reaction (including pain, redness and rash), joint pain, urinary tract infection, diarrhea, chest cold, pain in legs or arms and shortness of breath2,3.

Novartis has obtained global rights to develop, manufacture and commercialize Leqvio under a license and collaboration agreement with Alnylam Pharmaceuticals, a leader in RNAi therapeutics.


Ref : https://en.wikipedia.org/wiki/Inclisiran
https://www.bachem.com/news/galnac-delivering-promise-of-oligonucleotides/

Monday, March 1, 2021

FDA Approves Amondys 45 (casimersen) Injection for the Treatment of Duchenne Muscular Dystrophy (DMD) in Patients Amenable to Skipping Exon 45

In continuation of my update on antisense oligonucleotides 

Sarepta Therapeutics, Inc. the leader in precision genetic medicine for rare diseases, today announced that the U.S. Food and Drug Administration (FDA) has approved Amondys 45 (casimersen). Amondys 45 is an antisense oligonucleotide from Sarepta’s phosphorodiamidate morpholino oligomer (PMO) platform, indicated for the treatment of Duchenne muscular dystrophy (DMD) in patients with a confirmed mutation amenable to exon 45 skipping. This indication is based on a statistically significant increase in dystrophin production in skeletal muscle observed in patients treated with Amondys 45, which is reasonably likely to predict clinical benefit for those patients who are exon 45 amenable. Consistent with the accelerated approval pathway, the continued approval of Amondys 45 may be contingent on confirmation of a clinical benefit in confirmatory trials.

The ESSENCE trial – a placebo-controlled confirmatory trial to support the Amondys 45 approval – is ongoing and expected to conclude in 2024.

Although kidney toxicity was not observed in the clinical studies with Amondys 45, kidney toxicity, including potentially fatal glomerulonephritis, has been observed after administration of some antisense oligonucleotides. Kidney function should be monitored in patients taking Amondys 45. In the clinical trial, the most common adverse reactions observed in at least 20% of patients treated with Amondys 45 and at least 5% more frequently than in placebo were (Amondys 45, placebo): upper respiratory tract infections (65%, 55%), cough (33%, 26%), fever (33%, 23%), headache (32%, 19%), joint pain (21%, 10%), and pain in mouth and throat (21%, 7%).

“This is an important day for Sarepta and, far more importantly, for the patients that we serve. After years of scientific commitment, investment and development, the approval of Amondys 45, Sarepta’s third approved RNA therapy, offers treatment to the 8% of the DMD community who have a confirmed exon 45 amenable mutation,” said Doug Ingram, president and chief executive officer, Sarepta. “Along with our other approved RNA therapies, we can now offer treatment options for nearly 30% of Duchenne patients in the U.S. And our commitment to bring therapies to the greatest percentage of the DMD community as soon as possible continues.”

“Decades of research and commitment have fueled and now accelerate our progress towards new treatments for Duchenne,” said Marissa Penrod, founder of Team Joseph and parent of an 18-year old with Duchenne. “The extraordinary diligence and persistence of the Duchenne community – patients and families, clinicians and researchers – have led us to today’s approval, where we now have exon-skipping treatments for almost a third of those with Duchenne.”   


Tuesday, March 24, 2020

FDA Approves Vyondys 53 (golodirsen) Injection for the Treatment of Duchenne Muscular Dystrophy (DMD) in Patients Amenable to Skipping Exon 53

In continuation of my update on oligonucleotide.

VYONDYS 53 (golodirsen) Structural Formula - Illustration


Sarepta Therapeutics, Inc. (NASDAQ:SRPT), the leader in precision genetic medicine for rare diseases, today announced that the U.S. Food and Drug Administration (FDA) has approved Vyondys 53™ (golodirsen). Vyondys 53 is an antisense oligonucleotide from Sarepta’s phosphorodiamidate morpholino oligomer (PMO) platform, indicated for the treatment of Duchenne muscular dystrophy (DMD) in patients with a confirmed mutation amenable to exon 53 skipping. This indication is based on a statistically significant increase in dystrophin production in skeletal muscle observed in patients treated with Vyondys 53, which is reasonably likely to predict clinical benefit for those patients who are exon 53 amenable. Consistent with the accelerated approval pathway, the continued approval of Vyondys 53 may be contingent on confirmation of a clinical benefit in this post-marketing confirmatory trial.

Sarepta’s placebo-controlled, post-marketing confirmatory trial to support the Vyondys 53 accelerated approval – titled ESSENCE – is currently enrolling and expected to conclude by 2024.
Hypersensitivity reactions, including rash, pyrexia (fever), pruritis, urticaria (hives), dermatitis, and skin exfoliation have occurred in patients who were treated with Vyondys 53. Renal toxicity was observed in animal studies. Although not observed in the clinical studies with Vyondys 53, renal toxicity, including potentially fatal glomerulonephritis, has been observed after administration of some antisense oligonucleotides. The most common adverse reactions that occurred in at least 20% of Vyondys 53-treated patients and more frequently than in placebo-treated patients were headache (41%), pyrexia (41%), fall (29%), abdominal pain (27%), nasopharyngitis (27%), cough (27%), vomiting (27%), and nausea (20%).
Following a New Drug Application (NDA) submission to and review by the Division of Neurology Products (the Review Division) for Vyondys 53, which the Review Division recommended for approval, the Office of Drug Evaluation 1 issued a complete response letter (CRL) in August of 2019. Thereafter, Sarepta made a formal dispute resolution request as outlined in relevant FDA Guidance. With the support of the Review Division, the matters raised in the CRL were rapidly evaluated and resolved by Dr. Peter Stein, Director of the Office of New Drugs (OND). OND granted the Company’s appeal and Sarepta re-submitted its NDA to the Review Division, which worked expeditiously to review and approve Vyondys 53.
“Today is monumental for Sarepta and, more importantly, for the DMD community,” said Doug Ingram, president and chief executive officer, Sarepta. “Vyondys 53, our second approved exon-skipping RNA therapy for DMD, may treat up to 8% of the DMD community, representing those patients who have a confirmed exon 53 amenable mutation. Along with EXONDYS 51® (eteplirsen), we now offer treatment options for approximately 20% of those with DMD in the U.S.”
Ingram continued, “In the span of four months, we commenced and completed the formal dispute resolution process culminating in the grant of our appeal, resubmitted our NDA and obtained an approval – a great benefit to DMD patients awaiting treatment. This unprecedented timing could not have been achieved without the commitment of the Review Division under the leadership of Dr. Billy Dunn, and the Office of New Drugs, which expeditiously heard and granted our appeal. Along with the DMD community, we owe our gratitude to both the Review Division and the OND for their objective, evidence-based approach to this review, for their fairness, and for the sense of urgency with which they addressed and resolved the CRL and granted this approval.”
“With the approval of Vyondys 53, up to another 8% of Duchenne families will have a therapy to treat this devastating disease,” said Pat Furlong, founding president and chief executive officer, Parent Project Muscular Dystrophy (PPMD). “For 25 years, PPMD has been working with researchers, clinicians, industry, and the Duchenne community to find treatments for all people living with Duchenne. And while we need to ensure that these approved therapies are accessible for patients, today we celebrate this approval and thank Sarepta for their continued leadership in the fight to end Duchenne.”
Vyondys 53 is priced at parity to EXONDYS 51, the price of which has not increased since its launch in 2016. Patients and physicians can access more information at www.SareptAssist.com or by calling 1-888-727-3782.

About Vyondys 53

Vyondys 53 is an antisense oligonucleotide indicated for the treatment of Duchenne muscular dystrophy in patients who have a confirmed mutation of the DMD gene that is amenable to exon 53 skipping. Vyondys 53 uses Sarepta’s proprietary phosphorodiamidate morpholino oligomer (PMO) chemistry and exon-skipping technology to bind to exon 53 of dystrophin pre-mRNA, resulting in exclusion, or “skipping,” of this exon during mRNA processing in patients with genetic mutations that are amenable to exon 53 skipping. Exon skipping is intended to allow for production of an internally truncated dystrophin protein.
https://www.rxlist.com/vyondys-53-drug.htm


Wednesday, March 4, 2020

FDA Approves Givlaari (givosiran) for Acute Hepatic Porphyria


Alnylam Pharmaceuticals, Inc. (Nasdaq: ALNY), the leading RNAi therapeutics company, announced today that the U.S. Food and Drug Administration (FDA) approved Givlaari (givosiran) injection for subcutaneous use for the treatment of adults with acute hepatic porphyria (AHP). AHP is a family of ultra-rare, genetic diseases characterized by debilitating, potentially life-threatening attacks and, for some patients, chronic manifestations that negatively impact daily functioning and quality of life. Long-term complications of AHP can include chronic neuropathic pain, hypertension, chronic kidney disease and liver disease. Givlaari was shown to significantly reduce the rate of porphyria attacks that required hospitalizations, urgent healthcare visits or IV hemin administration at home.

“We believe the approval of Givlaari represents a landmark event for the advancement of precision genetic medicines, providing new hope for patients and their caregivers living with the debilitating manifestations of AHP and unpredictable nature of AHP attacks, as well as for the doctors who diagnose and treat these patients. We are grateful to the investigators, patients and families who have helped make this new treatment option a reality for the AHP community. We also commend the FDA for recognizing the immense medical need and granting this approval so quickly,” said John Maraganore, Ph.D., Chief Executive Officer of Alnylam. “Givlaari now becomes our second RNAi therapeutic to be approved in the last 16 months, and the world’s first-ever GalNAc-conjugate RNA therapeutic to be approved, representing a watershed moment for a technology uniquely pioneered by Alnylam scientists. We believe today’s news reinforces the promise and potential of RNAi therapeutics as a whole new class of medicines and brings us one important step closer to fulfilling our Alnylam 2020 goals of building a multi-product, global commercial company with a deep clinical pipeline to drive growth and an organic product engine to fuel sustainable innovation.”
The FDA approval of Givlaari was received in less than four months after acceptance of the NDA, and was based on positive results from the ENVISION Phase 3 study, a randomized, double-blind, placebo-controlled, multinational study of 94 patients with AHP, at 36 study sites in 18 countries – the largest ever interventional study conducted in AHP. In ENVISION, AHP patients on Givlaari experienced 70% (95% CI: 60%, 80%) fewer porphyria attacks compared to placebo. Givlaari also resulted in a similar reduction in intravenous hemin use, as well as reductions in urinary aminolevulinic acid (ALA), and urinary porphobilinogen (PBG).
In the pivotal ENVISION study, the most common adverse reactions (reported in at least 20% of patients) with Givlaari were nausea (27%) and injection site reactions (25%). Other adverse reactions seen in patients treated with Givlaari (occurring over 5% more frequently than placebo) include rash, serum creatinine increase, transaminase elevations and fatigue. As previously reported, one patient in the Givlaari clinical development program experienced an anaphylactic reaction which resolved with medical management.
“Adults with AHP now have a new treatment option that has demonstrated the ability to reduce the frequency of porphyria attacks by specifically addressing factors associated with attacks and other disease manifestations of AHP,” said Manisha Balwanii, M.D., M.S, Associate Professor of the Department of Genetics and Genomic Sciences and Department of Medicine at the Icahn School of Medicine at Mount Sinai and principal investigator of the ENVISION study. “With the approval of Givlaari, and based on the efficacy data from the ENVISION study, I hope to see my patients and those across the country be able to live more normal lives with fewer porphyria attacks.”
“The FDA approval of Givlaari is an important milestone for our community, as we now have a new treatment option for adults living with acute hepatic porphyria,” said Kristen Wheeden, Executive Director, American Porphyria Foundation. “AHP can have a profound impact on the lives of patients and their families. Porphyria attacks are associated with severe, incapacitating pain, often requiring hospitalization for management. In addition, many patients struggle on a daily basis with chronic symptoms related to their disease. The approval of Givlaari is exciting for our community.”
Alnylam is committed to helping people access the medicines they are prescribed and will be offering comprehensive support services for people prescribed Givlaari through Alnylam Assist®. Visit AlnylamAssist.com for more information or call 1-833-256-2748.
Givlaari is expected to be available for shipment to healthcare providers in the U.S. by year-end. HCPs can initiate the process now by visiting www.AlnylamAssist.com and completing and submitting a Start Form.
In August, Alnylam announced a U.S. gastrointestinal (GI) disease education and promotional agreement for Givlaari with Ironwood Pharmaceuticals, Inc., a GI healthcare company. Under the agreement, Alnylam will leverage Ironwood’s leading capabilities in GI to help raise awareness of AHP among gastroenterologists and other healthcare practitioners in the U.S. Ironwood will also participate in Givlaari promotional efforts, augmenting Alnylam’s broader commercialization activities.
https://en.wikipedia.org/wiki/Givosiran

Saturday, May 18, 2019

FDA Approves Dovato (dolutegravir/lamivudine) for HIV-1 Infection

In continuation of my update on Dolutegravir & Lamivudine


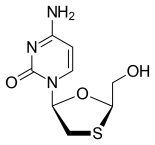
Dolutegravir.svg
Dolutegravir (DTG)

Lamivudine structure.svg
Lamivudine, commonly called 3TC

ViiV Healthcare  announced the US Food and Drug Administration (FDA) approval of Dovato, a complete, once-daily, single-tablet regimen of dolutegravir (DTG) 50 mg and lamivudine (3TC) 300 mg for the treatment of HIV-1 infection in adults with no antiretroviral (ARV) treatment history and with no known resistance to either DTG or 3TC. Dovato, a two-drug regimen (2DR), reduces exposure to the number of ARVs from the start of treatment, while still maintaining the efficacy and high barrier to resistance of a traditional DTG-based three-drug regimen.
Deborah Waterhouse, CEO, ViiV Healthcare, said: “Building on our innovative portfolio of medicines, Dovato is powered by dolutegravir, an antiretroviral included in multiple combination therapies and the most prescribed integrase inhibitor in the world, 2 coupled with the established profile of lamivudine. With Dovato, the first complete, single-tablet, two-drug regimen for treatment-naïve adults, ViiV Healthcare is delivering what patients are requesting—a chance to treat their HIV-1 infection with as few drugs as possible, marking a significant step in HIV treatment.”
The approval of Dovato is supported by the landmark global GEMINI 1 and 2 studies that included more than 1,400 HIV-1 infected adults. In these studies, DTG + 3TC demonstrated non-inferiority based on plasma HIV-1 RNA <50 copies per milliliter (c/mL), a standard measure of HIV-1 control, at Week 48 when compared to a three-drug regimen of DTG and two nucleoside reverse transcriptase inhibitors (NRTIs), tenofovir disoproxil fumarate/emtricitabine (TDF/FTC), in treatment-naïve, HIV-1 infected adults. The safety results for DTG + 3TC seen in GEMINI 1 and 2 were consistent with the product labelling for DTG and 3TC. No patient who experienced virologic failure in either treatment arm developed treatment-emergent resistance.
Pedro Cahn, principal investigator for the GEMINI study program said: “People are now living longer with HIV and will spend a lifetime taking drugs to suppress their virus. The approval of the fixed dose combination of dolutegravir and lamivudine, a complete, single-tablet, two-drug regimen, marks a pivotal moment in the treatment of HIV-1. Treatment-naïve people living with the virus have a powerful option that delivers non-inferior efficacy to a dolutegravir-based three-drug regimen, allowing them to take fewer ARVs and get and remain suppressed.”
Jeff Berry, Test Positive Aware Network (TPAN), said: “The approval of Dovato is a welcome paradigm shift, as it brings an innovative treatment approach to newly diagnosed adults with HIV-1. By exposing patients to fewer drugs at the start of treatment, the hope is to help address concerns arising from overall management of prolonged ARV therapy.”
DTG/3TC as a complete, once-daily, single-tablet, two-drug regimen for HIV-1 therapy is currently under review by the European Medicines Agency (EMA) and regulatory authorities in Canada, Australia, Switzerland, and South Africa and several additional submissions are planned throughout 2019.

About Dovato (dolutegravir/lamivudine)

Dovato is approved as a complete regimen for the treatment of HIV-1 infection in adults with no known antiretroviral treatment history and with no known substitutions associated with resistance to either dolutegravir or lamivudine. Dovato is a once-daily, single-tablet, two-drug regimen that combines the integrase strand transfer inhibitor (INSTI) dolutegravir (Tivicay, 50 mg) with the nucleoside analogue reverse transcriptase inhibitor (NRTI) lamivudine (Epivir, 300 mg).
Like a DTG-based three-drug regimen, Dovato uses only two drugs to inhibit the viral cycle at two different sites. INSTIs, like dolutegravir, inhibit HIV replication by preventing the viral DNA from integrating into the genetic material of human immune cells (T-cells). This step is essential in the HIV replication cycle and is also responsible for establishing chronic infection. Lamivudine is an NRTI that works by interfering with the conversion of viral RNA into DNA which in turn stops the virus from multiplying.

https://en.wikipedia.org/wiki/Dolutegravir
https://en.wikipedia.org/wiki/Lamivudine

Wednesday, October 17, 2018

FDA Approves Onpattro (patisiran) Targeted RNA-based Therapy for Polyneuropathy Caused by hATTR


The U.S. Food and Drug Administration today approved Onpattro (patisiran) infusion for the treatment of peripheral nerve disease (polyneuropathy) caused by hereditary transthyretin-mediated amyloidosis (hATTR) in adult patients. This is the first FDA-approved treatment for patients with polyneuropathy caused by hATTR, a rare, debilitating and often fatal genetic disease characterized by the buildup of abnormal amyloid protein in peripheral nerves, the heart and other organs. It is also the first FDA approval of a new class of drugs called small interfering ribonucleic acid (siRNA) treatment.

Patisiran.png


This approval is part of a broader wave of advances that allow us to treat disease by actually targeting the root cause, enabling us to arrest or reverse a condition, rather than only being able to slow its progression or treat its symptoms. In this case, the effects of the disease cause a degeneration of the nerves, which can manifest in pain, weakness and loss of mobility,” said FDA Commissioner Scott Gottlieb, M.D. “New technologies like RNA inhibitors, that alter the genetic drivers of a disease, have the potential to transform medicine, so we can better confront and even cure debilitating illnesses. We’re committed to advancing scientific principles that enable the efficient development and review of safe, effective and groundbreaking treatments that have the potential to change patients’ lives.”
RNA acts as a messenger within the body’s cells, carrying instructions from DNA for controlling the synthesis of proteins. RNA interference is a process that occurs naturally within our cells to block how certain genes are expressed. Since its discovery in 1998, scientists have used RNA interference as a tool to investigate gene function and its involvement in health and disease. Researchers at the National Institutes of Health, for example, have used robotic technologies to introduce siRNAs into human cells to individually turn off nearly 22,000 genes.
This new class of drugs, called siRNAs, work by silencing a portion of RNA involved in causing the disease. More specifically, Onpattro encases the siRNA into a lipid nanoparticle to deliver the drug directly into the liver, in an infusion treatment, to alter or halt the production of disease-causing proteins.
Affecting about 50,000 people worldwide, hATTR is a rare condition. It is characterized by the buildup of abnormal deposits of protein fibers called amyloid in the body's organs and tissues, interfering with their normal functioning. These protein deposits most frequently occur in the peripheral nervous system, which can result in a loss of sensation, pain, or immobility in the arms, legs, hands and feet. Amyloid deposits can also affect the functioning of the heart, kidneys, eyes and gastrointestinal tract. Treatment options have generally focused on symptom management.
Onpattro is designed to interfere with RNA production of an abnormal form of the protein transthyretin (TTR). By preventing the production of TTR, the drug can help reduce the accumulation of amyloid deposits in peripheral nerves, improving symptoms and helping patients better manage the condition.
“There has been a long-standing need for a treatment for hereditary transthyretin-mediated amyloidosis polyneuropathy. This unique targeted therapy offers these patients an innovative treatment for their symptoms that directly affects the underlying basis of this disease,” said Billy Dunn, M.D., director of the Division of Neurology Products in the FDA’s Center for Drug Evaluation and Research.
The efficacy of Onpattro was shown in a clinical trial involving 225 patients, 148 of whom were randomly assigned to receive an Onpattro infusion once every three weeks for 18 months, and 77 of whom were randomly assigned to receive a placebo infusion at the same frequency. The patients who received Onpattro had better outcomes on measures of polyneuropathy including muscle strength, sensation (pain, temperature, numbness), reflexes and autonomic symptoms (blood pressure, heart rate, digestion) compared to those receiving the placebo infusions. Onpattro-treated patients also scored better on assessments of walking, nutritional status and the ability to perform activities of daily living.
The most common adverse reactions reported by patients treated with Onpattro are infusion-related reactions including flushing, back pain, nausea, abdominal pain, dyspnea (difficulty breathing) and headache. All patients who participated in the clinical trials received premedication with a corticosteroid, acetaminophen, and antihistamines (H1 and H2 blockers) to reduce the occurrence of infusion-related reactions. Patients may also experience vision problems including dry eyes, blurred vision and eye floaters (vitreous floaters). Onpattro leads to a decrease in serum vitamin A levels, so patients should take a daily Vitamin A supplement at the recommended daily allowance.
The FDA granted this application Fast Track, Priority Review and Breakthrough Therapy designations. Onpattro also received Orphan Drug designation, which provides incentives to assist and encourage the development of drugs for rare diseases.

https://en.wikipedia.org/wiki/Patisiran
----------------------------------------------------------------------------------------


FDA Approves Onpattro (patisiran) Targeted RNA-based Therapy for Polyneuropathy Caused by hATTR

Thursday, January 4, 2018

Garlic compound can combat robust bacteria in patients with chronic infections

An active sulphurous compound found in garlic can be used to fight robust bacteria in patients with chronic infections, a new study from the University of Copenhagen indicates. Here the researchers show that the garlic compound is able to destroy important components in the bacteria's communication systems, which involve regulatory RNA molecules.
'We really believe this method can lead to treatment of patients, who otherwise have poor prospects. Because chronic infections like cystic fibrosis can be very robust. But now we, together with a private company, have enough knowledge to further develop the garlic drug and test it on patients', says Assistant Professor Tim Holm Jakobsen from the Costerton Biofilm Center at the Department of Immunology and Microbiology.
The study is the latest addition from a research group headed by Professor Michael Givskov, which since 2005 has focussed on garlic's effect on bacteria. At the time they learned that garlic extract is able to inhibit bacteria, and in 2012 they showed that the sulphurous compound ajoene found in garlic is responsible for the effect. The new study, which has been published in the scientific journal Scientific Reports, takes an even closer look and documents ajoene's ability to inhibit small regulatory RNA molecules in two types of bacteria.
'The two types of bacteria we have studied are very important. They are called Staphylococcus aureus and Pseudomonas aeruginosa. They actually belong to two very different bacteria families and are normally fought using different methods. But the garlic compound is able to fight both at once and therefore may prove an effective drug when used together with antibiotics', says Tim Holm Jakobsen.
Previous studies have shown that garlic appears to offer the most powerful, naturally occurring resistance to bacteria. In addition to inhibiting the bacteria's RNA molecules, the active garlic compound also damages the protective slimy matrix surrounding the bacteria, the so-called biofilm. When the biofilm is destroyed or weakened, both antibiotics and the body's own immune system are able to attack the bacteria more directly and thus remove the infection.
In 2012 the researchers took out a patent on the use of ajoene to fight bacterial infections. Now the company Neem Biotech has bought the licence to use the patent. Their medical product, NX-AS-401, which aims to treat patients with cystic fibrosis, has now obtained a so-called 'orphan drug designation'. This means that clinical trials on patients will be conducted soon.
If the clinical trials show good results, the drug can be marketed as the first in a series of antimicrobial connections with brand new modes of action developed by Givskov's research team.
Ref : http://healthsciences.ku.dk/news/2017/11/garlic/

Wednesday, November 22, 2017

FDA Approves Juluca, First Two-Drug Regimen for HIV Patients

In continuation of my updates on dolutegravir and rilpivirine,
The FDA has approved the first complete treatment regimen containing only two drugs to treat certain adults with human immunodeficiency virus type 1 (HIV-1) instead of the three or more drugs included in standard HIV treatment.

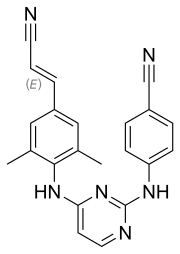
Juluca (dolutegravir/rilpivirine, ViiV Healthcare) is a fixed-dose tablet approved to treat adults with HIV-1 infections whose virus is currently suppressed (HIV-1 RNA less than 50 copies per mL) on a stable regimen for at least six months, with no history of treatment failure and no known substitutions associated with resistance to the individual components of the new combination. Dolutegravir 50 mg (ViiV Healthcare) is an integrase strand transfer inhibitor, and rilpivirine 25 mg (Janssen Therapeutics) is a non-nucleoside reverse transcriptase inhibitor.
Rilpivirine.svg rilpivirine    Dolutegravir.svg Dolutegravir
 “Limiting the number of drugs in any HIV treatment regimen can help reduce toxicity for patients,” said Debra Birnkrant, MD, director of the Division of Antiviral Products in the FDA’s Center for Drug Evaluation and Research.
HIV weakens a person’s immune system by destroying important cells that fight disease and infection. According to the Centers for Disease Control and Prevention, an estimated 1.1 million people in the United States are living with HIV, and the disease remains a significant cause of death for certain populations.
This FDA approval is based primarily on data from two pivotal phase 3 clinical trials, SWORD-12 and SWORD-2,2 which showed the two-drug regimen achieved non-inferior viral suppression (HIV-1 RNA less than 50 copies per mL) at 48 weeks compared with patients’ three- or four-drug current antiretroviral regimen (CAR) in both pooled and individual analyses of the SWORD-1 and SWORD-2 studies (dolutegravir/rilpivirine 486/513 [95%], CAR 485/511 [95%]; adjusted difference, –0.2%; 95% confidence interval, –3.0% to  2.5%, pooled analysis). Virological suppression rates were similar between treatment arms. Drug related adverse events and adverse events leading to withdrawal occurred in low frequencies in both arms of the study, but more frequently in the investigational arm.
The most common side effects in patients taking Juluca were diarrhea and headache. Serious side effects include skin rash and allergic reactions, liver problems, and depression or mood changes. Juluca should not be given with other anti-HIV drugs and may have drug interactions with other commonly used medications.
Ref : https://www.viivhealthcare.com/media/press-releases/2017/november/viiv-healthcare-announces-us-fda-approval-for-juluca.aspx