Thursday, September 3, 2026
Nuvalent Announces Submission of New Drug Application to FDA for Neladalkib in TKI Pre-treated Advanced ALK-Positive NSCLC
Friday, August 21, 2026
FDA Accepts New Drug Application for Zipalertinib for the Treatment of Non-Small Cell Lung Cancer
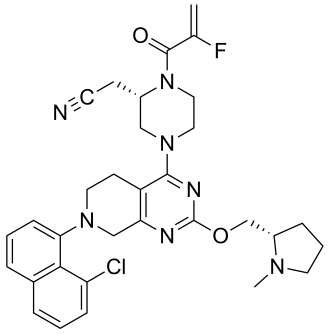
Taiho Oncology, Inc., Taiho Pharmaceutical Co., Ltd., and Cullinan Therapeutics, Inc.(Nasdaq: CGEM) announced the U.S. Food and Drug Administration (FDA) acceptance of New Drug Application (NDA) for zipalertinib for the treatment of patients with locally advanced or metastatic non-small cell lung cancer (NSCLC) with epidermal growth factor receptor (EGFR) exon 20 insertion (ex20ins) mutations whose disease has progressed on or after platinum-based chemotherapy, with or without amivantamab. The Prescription Drug User Fee Act (PDUFA) target action date is February 27, 2027.
- NDA submission based on the Phase 2b REZILIENT1 clinical trial, which demonstrated clinically meaningful and durable responses in patients with relapsed EGFR exon 20 insertion–mutated NSCLC
- Prescription Drug User Fee Act (PDUFA) target action date is February 27, 2027
The NDA is supported by data from the Phase 2b part of the REZILIENT1 clinical trial of zipalertinib monotherapy in patients with NSCLC harboring EGFR ex20ins mutations who have received prior therapy. The study met its primary endpoint of objective response rate. Study results from REZILIENT1 were presented at the 2025 American Society of Clinical Oncology (ASCO) Annual Meeting and simultaneously published in the Journal of Clinical Oncology.
“Zipalertinib was discovered at Taiho Pharmaceutical Co., Ltd., and has been developed with a focus on addressing the unmet needs of patients with EGFR exon 20 insertion-mutated non-small cell lung cancer,” said Harold Keer, MD, PhD, Chief Medical Officer, Taiho Oncology. “The FDA’s acceptance of the NDA for zipalertinib is an important milestone for this program, and we look forward to working with FDA during the review process.”
“Zipalertinib is a compound created using Taiho Pharmaceutical’s proprietary drug discovery and development technologies, Cysteinomix, with the aim of delivering a new treatment option to address high unmet medical needs,” said Takeshi Sagara, PhD, Executive Director, Board Member, Medical Affairs, Translational Development, Clinical Development, Discovery and Preclinical Research at Taiho Pharmaceutical. “The FDA’s acceptance of the NDA represents an important milestone, reflecting the scientific and clinical data accumulated to date. We will continue to work closely with Taiho Oncology, Cullinan Therapeutics and the FDA throughout the review process, with the shared goal of ultimately delivering a new treatment option to patients with non-small cell lung cancer EGFR exon 20 insertion mutations.”
“FDA acceptance of the zipalertinib NDA is an important step toward making zipalertinib available for people living with non-small cell lung cancer with EGFR exon 20 insertion mutations, who continue to face limited treatment options,” said Jeffrey Jones, MD, MBA, Chief Medical Officer, Cullinan Therapeutics. “We are deeply grateful to the patients and families who have participated in the REZILIENT program, and to the investigators, study teams, and advocates whose collaboration made achievement of this milestone possible. We believe zipalertinib has the potential to help address a significant unmet need, and we look forward to working with our partners at Taiho with the goal of bringing zipalertinib to patients waiting for new treatment options.”
Summary of Primary Study Results:
- Zipalertinib demonstrated clinically meaningful efficacy in the primary efficacy population (n=176), including 51 patients who had received prior amivantamab.
- The confirmed objective response rate (ORR) was 35%. Median duration of response (mDOR) was 8.8 months
- In patients treated after prior platinum-based chemotherapy only (n=125), ORR was 40% with a mDOR of 8.8 months.
- In exploratory subgroup analyses:
- Patients who had received prior amivantamab without other ex20ins-targeted therapy (n=30) showed a confirmed ORR of 30% and mDOR of 14.7 months.
- Patients with brain metastases (n=68) showed a confirmed ORR of 31% and a mDOR of 8.3 months.
- The safety profile of zipalertinib was manageable and consistent with previously reported data.¹ The most common treatment-emergent adverse events were paronychia, rash, anemia, dermatitis acneiform, diarrhea, dry skin, nausea and stomatitis. Most treatment-emergent adverse events were grade 1 or 2 per NCI-Common Terminology Criteria for Adverse Events (CTCAE v5.0).
Zipalertinib is an oral EGFR tyrosine kinase inhibitor. Zipalertinib received Breakthrough Therapy Designation in 2021 for the treatment of patients with locally advanced or metastatic NSCLC harboring EGFR ex20ins mutations who have previously received platinum‑based systemic chemotherapy.
Wednesday, August 5, 2026
Nuvalent Announces Submission of New Drug Application to FDA for Neladalkib in TKI Pre-treated Advanced ALK-Positive NSCLC
Saturday, July 25, 2026
FDA Grants Accelerated Approval to Hyrnuo (sevabertinib) for Non-Squamous Non-Small Cell Lung Cancer
Tuesday, December 16, 2025
FDA Grants Accelerated Approval to Hernexeos (zongertinib) for Previously Treated Patients with HER2-Mutant Advanced NSCLC
Monday, August 11, 2025
FDA Grants Accelerated Approval to Zegfrovy (sunvozertinib) for Non-Small Cell Lung Cancer with EGFR Exon 20 Insertion Mutations
Friday, July 25, 2025
FDA Approves Ibtrozi (taletrectinib) for Advanced ROS1-Positive Non-Small Cell Lung Cancer
Nuvation Bio Inc. announced the U.S. Food and Drug Administration (FDA) approval of Ibtrozi (taletrectinib) for the treatment of adult patients with locally advanced or metastatic ROS1-positive (ROS1+) non-small cell lung cancer (NSCLC). Ibtrozi is a highly selective, next-generation oral ROS1 tyrosine kinase inhibitor (TKI) designed to address some of the outstanding challenges of treating ROS1+ NSCLC. It has demonstrated high response rates with durable benefit and intracranial activity and is generally well tolerated, providing a new treatment option for patients with advanced ROS1+ NSCLC.
“The FDA approval of Ibtrozi marks a major milestone in the evolution of targeted therapy for advanced ROS1-positive NSCLC,” said David Hung, M.D., Founder, President and Chief Executive Officer of Nuvation Bio. “We believe one of the greatest threats to ROS1-positive lung cancer patients is disease progression, especially in the first-line setting. In pivotal trials, Ibtrozi delivered high response rates with sustained durability—truly meaningful benefits for patients. With its clinically proven efficacy and safety profile, we believe Ibtrozi has the potential to become a new standard for what targeted therapies can achieve in this type of lung cancer. With approvals for Ibtrozi now in the U.S. and China, and additional global filings underway, we remain committed to delivering innovative therapies that help patients stay ahead of their disease.”
ROS1+ NSCLC is a rare and aggressive form of lung cancer, accounting for approximately 2% of new NSCLC cases, or about 3,000 new diagnoses of advanced disease annually in the U.S. The median age at diagnosis for patients with this type of lung cancer is approximately 50 years old, and the disease is more likely to occur in people who have never smoked. Brain metastases are common and a leading cause of disease progression and mortality in this population.
“For people living with advanced ROS1-positive lung cancer, who tend to be diagnosed at a younger age, having another treatment option can make a real difference for them and their loved ones,” said Janet Freeman-Daily, Co-Founder and President of The ROS1ders. “The approval of this new targeted therapy is a meaningful step forward for the advanced ROS1+ lung cancer community and offers hope for patients facing the added challenge of cancer spreading to the brain.”
The FDA approval of Ibtrozi is supported by one of the largest global clinical trial programs in ROS1+ NSCLC to date, with over 300 patients enrolled in the pivotal TRUST-I and TRUST-II studies.
In TRUST-I, Ibtrozi achieved a confirmed overall response rate (cORR) of 90% in TKI-naïve patients. These findings were reinforced by the TRUST-II results, with a cORR of 85% in TKI-naïve patients. The median duration of response (DOR) was not yet reached for either trial, based on a cutoff date that is nearly five months later than that of the pooled TRUST-I and TRUST-II analysis published in April in the Journal of Clinical Oncology. For TRUST-I, with a median follow-up for responses of 40 months, the longest DOR was observed at 46.9 months and ongoing. For TRUST-II, with a median follow-up for responses of 19 months, the longest DOR was observed at 30.4 months and ongoing as of October 2024. Given the single-arm nature of the TRUST clinical studies, median progression-free survival (PFS) is not provided in the label.
Across the pivotal studies, consistent results were also observed among patients who were previously treated with a ROS1 TKI (TKI-pretreated). In TRUST-I, treatment with Ibtrozi achieved a cORR of 52% and median DOR of 13.2 months for TKI-pretreated patients, with median follow-up for responses of 33 months. In TRUST-II, treatment with Ibtrozi achieved a cORR of 62%, and as of October 2024 the median DOR was 19.4 months in these patients, with a median follow-up for responses of 19 months.
Brain metastases are among the most common and devastating complications in advanced ROS1+ NSCLC. Ibtrozi was designed to penetrate the central nervous system (CNS) and has demonstrated consistent intracranial responses in patients with measurable brain metastases at baseline. An intracranial response was achieved in 73% of TKI-naive patients (11/15) and 63% of TKI-pretreated patients (15/24).
“Patients living with advanced ROS1+ non-small cell lung cancer and their healthcare providers are in need of new treatment options,” added Nathan Pennell, M.D., Ph.D., TRUST study investigator and Professor of Medicine at the Cleveland Clinic. “Ibtrozi’s durability of response and ability to effectively penetrate the brain, coupled with a well-characterized and manageable safety profile, further addresses these critical needs for patients. I believe this now-approved therapy offers providers and patients a promising new option for the treatment of advanced ROS1+ non-small cell lung cancer.” Dr. Pennell is a compensated member of Nuvation Bio’s advisory committee.
Ibtrozi was generally well-tolerated, with most adverse events being low grade, transient and manageable. Patients infrequently (7%) discontinued treatment due to treatment-emergent adverse events (TEAEs). The most common adverse reactions (≥20%) included diarrhea (64%), nausea (47%), vomiting (43%), dizziness (22%), rash (22%), constipation (21%), and fatigue (20%). Overall, the majority of CNS events were mild to moderate (~90%) and resolved within days, and dose modifications due to these events were low (~5%). Approximately 90% of reported cases of dizziness were Grade 1 (mild) and transient. Liver enzyme elevations (AST 87%/ALT 85%) and QT prolongation (19%) were manageable with standard monitoring and dose modifications. Ibtrozi is approved as a 600 mg once-daily oral dose, supported by a half-life of approximately 66 hours and broad tissue distribution, including the brain, enabling sustained systemic and CNS exposure.
Saturday, August 24, 2024
FDA approves tepotinib for metastatic non-small cell lung cancer | FDA
Monday, August 12, 2024
FDA Approves Augtyro (repotrectinib) for the Treatment of Locally Advanced or Metastatic ROS1-Positive Non-Small Cell Lung Cancer (NSCLC)
Bristol Myers Squibb (NYSE: BMY) announced the U.S. Food and Drug Administration (FDA) approval of Augtyro (repotrectinib) for the treatment of adult patients with locally advanced or metastatic ROS1-positive non-small cell lung cancer (NSCLC). Administered as an oral therapy, Augtyro is a tyrosine kinase inhibitor (TKI) targeting ROS1 oncogenic fusions.
The approval is based on the TRIDENT-1 study, an open-label, single-arm, Phase 1/2 trial that evaluated Augtyro in TKI-naïve and TKI-pretreated patients.2 In TKI-naïve patients (n=71), the primary endpoint of objective response rate (ORR), defined as the percentage of people treated within a certain period of time whose tumor size decreased (partial response) or who no longer have signs of cancer (complete response),was 79% (95% Confidence Interval [CI]: 68 to 88).1,3 The median duration of response (mDOR) was 34.1 months. Among patients pretreated with one prior ROS1 TKI and no prior chemotherapy (n=56), the ORR was 38% (95% CI: 25 to 52) and the mDOR was 14.8 months.1 Among those who had measurable central nervous system (CNS) metastases at baseline, responses in intracranial lesions were observed in 7 of 8 TKI-naïve patients (n=71) and 5 of 12 of those who were TKI-pretreated (n=56).
“New treatment options continue to be needed for patients with ROS1 fusion-positive NSCLC that support important clinical goals, including achieving durable therapeutic responses,” said Jessica J. Lin, MD, TRIDENT-1 primary investigator and attending physician at the Center for Thoracic Cancers at Massachusetts General Hospital and Assistant Professor of Medicine at Harvard Medical School.4,5,6,7 “Based on the data we have seen in the TRIDENT-1 trial, repotrectinib has the potential to become a new standard of care option for patients with locally advanced or metastatic ROS1 fusion-positive lung cancer.”1
Augtyro is associated with the following Warnings & Precautions: central nervous system (CNS) effects, interstitial lung disease (ILD)/pneumonitis, hepatotoxicity, myalgia with creatine phosphokinase elevation, hyperuricemia, skeletal fractures, and embryo-fetal toxicity.1 Please see Important Safety Information below.
“While progress has been made in the treatment of NSCLC over the past decade, there is still a need to address this particularly difficult-to-treat form of the disease with innovative science and a targeted approach,” said Samit Hirawat, MD, executive vice president, chief medical officer, Global Drug Development, Bristol Myers Squibb.6,7 “As the only approved next-generation TKI for ROS1-positiveNSCLCpatients, Augtyro builds on our legacy of delivering transformational therapies for patients with thoracic cancers.”6,8,9
“ROS1-positive NSCLC patients and their families face a stressful journey because our cancer can be difficult to treat, especially when it spreads to the brain,” said Janet Freeman-Daily, co-founder and president of The ROS1ders, a patient advocacy organization.10 “Today’s approval brings a new treatment option for the ROS1-positive patient community, which gives us hope for more time with loved ones.”
Augtyro is designed to minimize interactions that can lead to certain forms of treatment resistance in ROS1-positive metastatic NSCLC patients. Itis expected to be available to patients in the U.S. in mid-December 2023. Bristol Myers Squibb thanks the patients and investigators involved in the TRIDENT-1 clinical trial program.
ref ;https://en.wikipedia.org/wiki/Repotrectinib
Thursday, January 19, 2023
FDA Approves Krazati (adagrasib) for Locally Advanced or Metastatic Non-Small Cell Lung Cancer (NSCLC) with a KRASG12C Mutation
This indication is approved under accelerated approval based on objective response rate (ORR) and duration of response (DOR). Continued approval for this indication may be contingent upon verification and description of a clinical benefit in a confirmatory trial(s).
"The FDA approval of Krazati is a positive development for thousands of patients with KRASG12C mutations, including the approximately 14% of patients with NSCLC adenocarcinomas histology that harbor a KRASG12C mutation.1 Mirati is thrilled to make Krazati available in a tablet formulation to patients in the U.S. with advanced NSCLC who have progressed beyond a first-line treatment for the historically difficult-to-treat KRAS mutation," David Meek, chief executive officer, Mirati Therapeutics, Inc., continued, "We look forward to continuing to advance our Krazati development program including several monotherapy and combination studies in KRASG12C-mutated solid tumors."
Krazati has demonstrated a positive benefit-risk profile with accelerated approval based on the Phase 2 registration-enabling cohort of the KRYSTAL-1 study, evaluating Krazati 600 mg capsules administered orally twice daily in 116 patients with KRASG12C-mutated advanced NSCLC who previously received treatment with a platinum-based regimen and an immune checkpoint inhibitor. The primary efficacy endpoints were confirmed ORR and DOR as evaluated by blinded independent central review (BICR) according to response evaluation criteria in solid tumors (RECIST v1.1).
The trial demonstrated an ORR of 43% (95% CI: 34-53) with 80% (95% CI: 71-87) of patients achieving disease control. The median DOR was 8.5 months (95% CI: 6.2-13.8).
In a pooled efficacy analysis (n=132) including Phase 1/1b NSCLC and registrational Phase 2 NSCLC cohorts from the KRYSTAL-1 study evaluating adagrasib as a single agent at 600 mg capsules orally twice daily, adagrasib showed an ORR of 44% and a disease control rate of 81% based on BICR, a median DOR of 12.5 months (95% CI, 7.3-NE) and median overall survival of 14.1 months (94% CI, 9.2-19.2).
The safety profile of Krazati was evaluated in a pooled patient population with NSCLC and other solid tumors as a single agent at 600 mg orally twice daily in 366 patients enrolled in KRYSTAL-1 and KRYSTAL-12. The most common (≥ 25%) adverse reactions were nausea, diarrhea, vomiting, fatigue, musculoskeletal pain, hepatotoxicity, renal impairment, edema, dyspnea and decreased appetite. Permanent discontinuation of Krazati due to an adverse reaction occurred in 13% of patients.
Although KRASG12C is the most common KRAS mutation in NSCLC, patients have had limited options for the treatment of this debilitating and difficult-to-treat condition.2,3
"The approval of Krazati offers an effective therapy for patients with advanced NSCLC harboring the KRASG12C mutation. The positive ORR and DOR results, as observed in previously treated patients with NSCLC harboring the KRASG12C mutation, demonstrate the effectiveness of Krazati as an option for these difficult-to-treat patients," said Shirish M. Gadgeel, MD, chief of the Division of Hematology and Oncology, Department of Internal Medicine, Henry Ford Cancer Institute/Henry Ford Health System.
"KRASG12C in NSCLC is an area of high unmet need and new treatment options offer patients and our community new hope for survivorship," said Bonnie J. Addario, co-founder and board chair of the GO2 Foundation for Lung Cancer. "I'm pleased that patients have options, there's more awareness of this disease and we are all focused on improving the journeys of people living with KRASG12C-mutated NSCLC."
The Company partnered with Agilent and QIAGEN to develop blood- and tissue-based companion diagnostics (CDx), respectively, for Krazati that are now available. With tissue and blood modalities for companion diagnostics, patients have more flexibility, and clinicians have greater options for biomarker testing. These solutions help to personalize a patient's treatment path.
Mirati Therapeutics is launching Mirati & Me, a comprehensive program dedicated to supporting patients, caregivers and the oncology community including coverage and access, financial, educational and emotional support services.
Thursday, January 27, 2022
Spectrum Pharmaceuticals Submits New Drug Application for Poziotinib
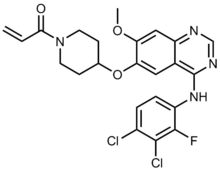
Spectrum Pharmaceuticals, a biopharmaceutical company focused on novel and targeted oncology therapies, announced the submission of its New Drug Application (NDA) for poziotinib to the U.S. Food and Drug Administration (FDA) for use in patients with previously treated locally advanced or metastatic non-small cell lung cancer (NSCLC) with HER2 exon 20 insertion mutations. The NDA submission is based on the positive results of Cohort 2 from the ZENITH20 clinical trial, which assessed the safety and efficacy of poziotinib. The product has received Fast Track designation and there is currently no treatment specifically approved by the FDA for this indication.
“The NDA submission for poziotinib marks an important step in achieving a first treatment for patients with HER2 exon 20 insertion mutations in lung cancer,” said Joe Turgeon, President and CEO of Spectrum Pharmaceuticals. “I want to thank the patients, investigators and our internal staff who have passionately worked to achieve this important milestone in an area of high unmet medical need.”
ZENITH20 Cohort 2 Clinical Results Summary
Results for Cohort 2 of the ZENITH20 clinical trial have been published in the Journal of Clinical Oncology (November 29, 2021), and can be accessed by clicking here.
Cohort 2 enrolled 90 patients who received an oral once daily dose of 16 mg of poziotinib. The intent-to-treat analysis demonstrated a confirmed objective response rate (ORR) of 27.8% (95% Confidence Interval (CI), 18.9%-38.2%). The observed lower bound of 18.9% exceeded the pre-specified lower bound of 17%. The median duration of response was 5.1 months and the median progression free survival was 5.5 months. In this cohort, 87% of patients had drug interruptions with 11 patients (12%) permanently discontinuing due to adverse events. 13 patients (14%) had treatment-related serious adverse events. As previously announced, the company had a successful pre-NDA meeting with the FDA which resulted in an agreement to submit an NDA for poziotinib. During the meeting, Spectrum confirmed with the FDA that Cohort 2 data could serve as the basis of an NDA submission. The company will continue to work with the FDA as appropriate, while the agency conducts its review.
More : https://investor.sppirx.com/news-releases/news-release-details/spectrum-pharmaceuticals-submits-new-drug-application-poziotinib
https://en.wikipedia.org/wiki/Poziotinib
Thursday, January 6, 2022
Spectrum Pharmaceuticals Submits New Drug Application for Poziotinib for metastatic NSCLC with HER2 exon 20 insertion mutations.
Spectrum Pharmaceuticals, a biopharmaceutical company focused on novel and targeted oncology therapies, announced that it has submitted its New Drug Application (NDA) for poziotinib to the U.S. Food and Drug Administration (FDA) for use in patients with previously treated locally advanced or metastatic non-small cell lung cancer (NSCLC) with HER2 exon 20 insertion mutations. The NDA submission is based on the positive results of Cohort 2 from the ZENITH20 clinical trial, which assessed the safety and efficacy of poziotinib. The product has received Fast Track designation and there is currently no treatment specifically approved by the FDA for this indication.
“The NDA submission for poziotinib marks an important step in achieving a first treatment for patients with HER2 exon 20 insertion mutations in lung cancer,” said Joe Turgeon, President and CEO of Spectrum Pharmaceuticals. “I want to thank the patients, investigators and our internal staff who have passionately worked to achieve this important milestone in an area of high unmet medical need.”
ZENITH20 Cohort 2 Clinical Results Summary
Results for Cohort 2 of the ZENITH20 clinical trial have been published in the Journal of Clinical Oncology (November 29, 2021), and can be accessed by clicking here.
Cohort 2 enrolled 90 patients who received an oral once daily dose of 16 mg of poziotinib. The intent-to-treat analysis demonstrated a confirmed objective response rate (ORR) of 27.8% (95% Confidence Interval (CI), 18.9%-38.2%). The observed lower bound of 18.9% exceeded the pre-specified lower bound of 17%. The median duration of response was 5.1 months and the median progression free survival was 5.5 months. In this cohort, 87% of patients had drug interruptions with 11 patients (12%) permanently discontinuing due to adverse events. 13 patients (14%) had treatment-related serious adverse events. As previously announced, the company had a successful pre-NDA meeting with the FDA which resulted in an agreement to submit an NDA for poziotinib. During the meeting, Spectrum confirmed with the FDA that Cohort 2 data could serve as the basis of an NDA submission. The company will continue to work with the FDA as appropriate, while the agency conducts its review.
About the ZENITH20 Clinical Trial
The ZENITH20 study consists of seven cohorts of NSCLC patients. Cohorts 1 (EGFR) and 2 (HER2) in previously treated NSCLC patients with exon 20 mutations and Cohort 3 (EGFR) in first-line patients have completed enrollment. Cohort 4 (HER2) in first-line NSCLC patients with exon 20 mutations is still enrolling patients. Cohorts 1- 4 are each independently powered for a pre-specified statistical hypothesis and the primary endpoint is objective response rate (ORR). Cohort 5 includes previously treated or treatment-naïve NSCLC patients with EGFR or HER2 exon 20 insertion mutations. Cohort 6 includes NSCLC patients with classical EGFR mutations who progressed while on treatment with first-line osimertinib and developed an additional EGFR mutation. Cohort 7 includes NSCLC patients with a variety of less common mutations in EGFR or HER2 exons 18-21 or the extracellular or transmembrane domains
https://en.wikipedia.org/wiki/Poziotinib
Friday, December 17, 2021
New Drug Application of Plinabulin (Response Letter from the FDA) for Prevention of Chemotherapy-Induced Neutropenia (CIN)...
BeyondSpring Pharmaceuticals announced the receipt of a Complete Response Letter (CRL) from the U.S. Food and Drug Administration (FDA) for the New Drug Application (NDA) seeking approval of plinabulin in combination with granulocyte colony-stimulating factor (G-CSF) for the prevention of chemotherapy-induced neutropenia (CIN). The FDA issued the CRL to indicate that they have completed their review of the application and have determined that it cannot be approved in its present form.
The FDA’s CRL indicated that the results of the single registrational trial (106 Phase 3) was not sufficiently robust to demonstrate benefit and that a second well controlled trial would be required to satisfy the substantial evidence requirement to support the CIN indication.
“BeyondSpring strongly believes that plinabulin in combination with G-CSF has significant potential to raise the standard of care in CIN, a devastating side effect of chemotherapy,” said Dr. Lan Huang, BeyondSpring’s co-founder, chief executive officer and chairwoman. “The Company plans to request a meeting with the FDA and remains committed to its goal of bringing plinabulin to cancer patients in need globally.”
BeyondSpring remains confident in the efficacy and safety data for plinabulin in combination with G-CSF for the prevention of CIN. The Company expects to work closely with the FDA to consider the possible future clinical pathway for CIN, which may include a second study.
Plinabulin is the first drug candidate submitted for FDA approval that has the potential to work in the critical first week of chemotherapy treatment before G-CSF is effective, to prevent the onset and improve clinical outcomes of CIN.
Plinabulin, BeyondSpring’s lead asset, is a selective immunomodulating microtubule-binding agent (SIMBA), which is a potent antigen presenting cell (APC) inducer. It is a novel, intravenous infused, patent-protected, NDA-stage asset for CIN prevention and a Phase 3 anti-cancer candidate for non-small cell lung cancer (NSCLC) with recently released positive topline data. Plinabulin triggers the release of the immune defense protein, GEF-H1, which leads to two distinct effects: first is a durable anticancer benefit due to the maturation of dendritic cells resulting in the activation of tumor antigen-specific T-cells to target cancer cells, and the second is early-onset of action in CIN prevention after chemotherapy by boosting the number of hematopoietic stem/progenitor cells (HSPCs). Plinabulin received Breakthrough Therapy designation and priority review from both U.S. and China FDA for the CIN prevention indication. As a “pipeline in a drug,” plinabulin is being broadly studied in combination with various immuno-oncology agents that could boost the effects of the PD-1/PD-L1 antibodies and re-sensitize PD-1/PD-L1 antibody-resistant patients.
https://en.wikipedia.org/wiki/Plinabulin
Thursday, December 2, 2021
FDA Approves Exkivity (mobocertinib) for EGFR Exon20 Insertion+ Non-Small Cell Lung Cancer (NSCLC)
Takeda Pharmaceutical Company Limited announced the U.S. Food and Drug Administration (FDA) approval of Exkivity (mobocertinib) for the treatment of adult patients with locally advanced or metastatic non-small cell lung cancer (NSCLC) with epidermal growth factor receptor (EGFR) exon 20 insertion mutations as detected by an FDA-approved test, whose disease has progressed on or after platinum-based chemotherapy. Exkivity, which was granted priority review and received Breakthrough Therapy Designation, Fast Track Designation and Orphan Drug Designation from the FDA, is the first and only approved oral therapy specifically designed to target EGFR Exon20 insertion mutations. This indication is approved under Accelerated Approval based on overall response rate (ORR) and DoR. Continued approval for this indication may be contingent upon verification and description of clinical benefit in a confirmatory trial.
“The approval of Exkivity introduces a new and effective treatment option for patients with EGFR Exon20 insertion+ NSCLC, fulfilling an urgent need for this difficult-to-treat cancer,” said Teresa Bitetti, president, Global Oncology Business Unit, Takeda. “Exkivity is the first and only oral therapy specifically designed to target EGFR Exon20 insertions, and we are particularly encouraged by the duration of the responses observed with a median of approximately 1.5 years. This approval milestone reinforces our commitment to meeting the needs of underserved patient populations within the oncology community.”
The FDA simultaneously approved Thermo Fisher Scientific’s Oncomine Dx Target Test as an NGS companion diagnostic for Exkivity to identify NSCLC patients with EGFR Exon20 insertions. NGS testing is critical for these patients, as it can enable more accurate diagnoses compared to polymerase chain reaction (PCR) testing, which detects less than 50% of EGFR Exon20 insertions.
“EGFR Exon20 insertion+ NSCLC is an underserved cancer that we have been unable to target effectively with traditional EGFR TKIs,” said Pasi A. Jänne, MD, PhD, Dana Farber Cancer Institute. “The approval of Exkivity (mobocertinib) marks another important step forward that provides physicians and their patients with a new targeted oral therapy specifically designed for this patient population that has shown clinically meaningful and sustained responses.”
“Patients with EGFR Exon20 insertion+ NSCLC have historically faced a unique set of challenges living with a very rare lung cancer that is not only underdiagnosed, but also lacking targeted treatment options that can improve response rates,” said Marcia Horn, executive director, Exon 20 Group at ICAN, International Cancer Advocacy Network. “As a patient advocate working with EGFR Exon20 insertion+ NSCLC patients and their families every day for nearly five years, I am thrilled to witness continued progress in the fight against this devastating disease and am grateful for the patients, families, healthcare professionals and scientists across the globe who contributed to the approval of this promising targeted therapy.”
The FDA approval is based on results from the platinum-pretreated population in the Phase 1/2 trial of Exkivity, which consisted of 114 patients with EGFR Exon20 insertion+ NSCLC who received prior platinum-based therapy and were treated at the 160 mg dose. Results were presented at the 2021 American Society of Clinical Oncology (ASCO) Annual Meeting from the Phase 1/2 trial and demonstrated a confirmed ORR of 28% per independent review committee (IRC) (35% per investigator) as well as a median DoR of 17.5 months per IRC, a median overall survival (OS) of 24 months and a median progression-free survival (PFS) of 7.3 months per IRC.