Showing posts with label New Drug Application. Show all posts

Thursday, September 3, 2026

Nuvalent Announces Submission of New Drug Application to FDA for Neladalkib in TKI Pre-treated Advanced ALK-Positive NSCLC


Nuvalent, Inc. (Nasdaq: NUVL), a clinical-stage biopharmaceutical company focused on creating precisely targeted therapies for clinically proven kinase targets in cancer, announced the submission to the U.S. Food and Drug Administration (FDA) of the Company's NDA for neladalkib, an investigational ALK-selective inhibitor, in TKI pre-treated advanced ALK-positive NSCLC.

New Drug Application (NDA) based on data in TKI pre-treated patients from the global ALKOVE-1 Phase 1/2 clinical trial
"The advancement of neladalkib from first clinical trial initiation to NDA submission in less than four years represents a remarkable pace in oncology drug development, underscoring the vigor and urgency our team brought to this program and our deep commitment to the ALK-positive NSCLC community," said Darlene Noci, A.L.M., Chief Development Officer at Nuvalent. "We would like to extend our sincere gratitude to the patients, families and investigators who have made this progress possible, and are committed to working closely with the FDA throughout the NDA review process toward our goal of bringing neladalkib to patients as quickly as possible."

The application is based on data in TKI pre-treated patients with advanced ALK-positive NSCLC treated with neladalkib in the global, registration-directed ALKOVE-1 Phase 1/2 clinical trial. In this population, neladalkib demonstrated encouraging overall activity, including intracranial responses, the ability to address key drivers of disease progression, and a generally well-tolerated safety profile consistent with its ALK-selective, TRK-sparing design. The company plans to share detailed results at a future medical meeting.

Neladalkib has received breakthrough therapy designation from the FDA for the treatment of patients with locally advanced or metastatic ALK-positive NSCLC who have been previously treated with 2 or more ALK tyrosine kinase inhibitors and orphan drug designation for ALK-positive NSCLC.

About Neladalkib
Neladalkib is an investigational, brain-penetrant, ALK-selective inhibitor created with the aim to overcome limitations observed with currently available ALK inhibitors. Neladalkib is designed to remain active in tumors that have developed resistance to first-, second-, and third-generation ALK inhibitors, including tumors with single or compound treatment-emergent ALK mutations such as G1202R. In addition, neladalkib is designed for central nervous system (CNS) penetrance to improve treatment options for patients with brain metastases, and to avoid inhibition of the structurally related tropomyosin receptor kinase (TRK) family. Together, these characteristics have the potential to avoid TRK-related CNS adverse events seen with dual TRK/ALK inhibitors and to drive deep, durable responses for patients across all lines of therapy. Neladalkib has received breakthrough therapy designation from the U.S. Food and Drug Administration (FDA) for the treatment of patients with locally advanced or metastatic ALK-positive non-small cell lung cancer (NSCLC) who have been previously treated with 2 or more ALK tyrosine kinase inhibitors and orphan drug designation for ALK-positive NSCLC.

About the ALKOVE-1 Phase 1/2 Clinical Trial
The ALKOVE-1 trial (NCT05384626) is a first-in-human Phase 1/2 clinical trial for patients with advanced ALK-positive NSCLC and other solid tumors. The completed Phase 1 portion enrolled ALK-positive NSCLC patients who previously received at least one ALK TKI, or patients with other ALK-positive solid tumors who had been previously treated or for whom no satisfactory standard of care exists. The Phase 1 portion of the trial was designed to evaluate the overall safety and tolerability of neladalkib, with additional objectives including determination of the recommended Phase 2 dose (RP2D), characterization of the pharmacokinetic profile, and evaluation of preliminary anti-tumor activity. The global, single arm, open label Phase 2 portion is designed with registrational intent for TKI pre-treated patients with advanced ALK-positive NSCLC. Global enrollment in ALKOVE-1 remains ongoing for adult and adolescent patients with ALK-positive solid tumors outside of NSCLC, and adolescent patients with ALK-positive NSCLC.

Nuvalent Announces Submission of New Drug Application to FDA for Neladalkib in TKI Pre-treated Advanced ALK-Positive NSCLC

Monday, August 24, 2026

Immunome Announces Submission of New Drug Application to U.S. FDA for Varegacestat for the Treatment of Adults with Desmoid Tumors


Immunome, Inc. (Nasdaq: IMNM), a biotechnology company committed to developing first-in-class and best-in-class targeted cancer therapies,  announced the submission of a New Drug Application (NDA) to the U.S. Food and Drug Administration (FDA) for varegacestat, an investigational, oral, once-daily gamma secretase inhibitor (GSI), for the treatment of adults with desmoid tumors.

  • New Drug Application (NDA) supported by positive Phase 3 RINGSIDE results, including significant improvement in progression-free survival vs. placebo (hazard ratio = 0.16, p<0.0001)
  • Trial also met all key secondary endpoints, with varegacestat delivering an objective response rate of 56%
  • Detailed Phase 3 RINGSIDE data selected for oral presentation at the 2026 ASCO Annual Meeting

“The varegacestat NDA submission marks an important milestone for Immunome. It reflects the strength of the RINGSIDE dataset and the commitment of the team advancing this program,” said Clay B. Siegall, Ph.D., President and Chief Executive Officer of Immunome. “We believe varegacestat has the potential to provide adults with progressing desmoid tumors with a meaningful new oral treatment option, and we are grateful to the patients, families, investigators and study site teams whose participation made this submission possible.”

The NDA is supported by positive results from the global, randomized, double-blind, placebo-controlled Phase 3 RINGSIDE trial of varegacestat in patients with progressing desmoid tumors.

The trial met its primary endpoint of improving progression-free survival, demonstrating a statistically significant and clinically meaningful improvement vs. placebo, with an 84% reduction in the risk of disease progression or death (hazard ratio (HR) = 0.16, 95% CI: 0.071, 0.375; p<0.0001). The confirmed objective response rate (ORR) based on RECIST v1.1 was 56% with varegacestat vs. 9% with placebo (p<0.0001), as assessed by blinded independent central review.

In an exploratory analysis, varegacestat demonstrated a median best change in tumor volume of -83% vs. +11% with placebo, as assessed by blinded independent central review. In addition, the trial met all key secondary endpoints, with varegacestat achieving statistically significant improvements vs. placebo in landmark tumor volume reduction and worst pain intensity.

Varegacestat was generally well tolerated, with a manageable safety profile consistent with the GSI class. The most common adverse events for participants in the treatment arm were diarrhea (82%), fatigue (44%), rash (43%), nausea (35%) and cough (34%). Most events were grade 1 or 2.

Immunome previously announced that data from RINGSIDE has been selected for presentation in an oral abstract session at the 2026 American Society of Clinical Oncology (ASCO) Annual Meeting, taking place May 29-June 2, 2026 in Chicago.

About the RINGSIDE Trial

The global, randomized, double-blind, placebo-controlled Phase 3 RINGSIDE trial (NCT04871282) evaluated the efficacy and safety of varegacestat in patients with progressing desmoid tumors. A total of 156 patients were randomized to receive varegacestat 1.2 mg daily or placebo until disease progression or death, representing the largest randomized study in this population. The primary endpoint of the trial was progression-free survival as assessed by blinded independent central review. Statistically controlled secondary endpoints were confirmed ORR using RECIST v1.1 and change in tumor volume at week 24, both determined by blinded independent central review, as well as change in pain intensity at week 12 as determined using a patient reported outcome instrument. Additional secondary endpoints included duration of response, best reduction in tumor volume, patient-reported outcomes, and safety and tolerability. RINGSIDE includes an open-label extension phase, which is ongoing.

About Desmoid Tumors

Desmoid tumors (also known as aggressive fibromatosis or desmoid-type fibromatosis) are aggressive non-metastatic soft tissue tumors that are prone to recurrence. Approximately 1,000-1,650 people are diagnosed with desmoid tumors each year in the United States, and there are approximately 10,000-11,000 actively managed patients. Those affected face debilitating pain, deformity and, in some cases, life-threatening organ damage. The chronic pain and physical limitations associated with desmoid tumors lead to a high clinical burden and impaired quality of life. Although desmoid tumors are not considered cancerous, they often require systemic treatment to prevent permanent disability and alleviate disease burden.

About Varegacestat

Varegacestat (formerly AL102) is an investigational, oral, once-daily gamma secretase inhibitor. In December 2025, Immunome reported positive topline results for the Phase 3 RINGSIDE trial of varegacestat in adults with progressing desmoid tumors. Data from RINGSIDE have been selected for oral presentation at the 2026 ASCO Annual Meeting.

https://www.medkoo.com/products/7653


Friday, August 14, 2026

FDA Accepts New Drug Application for Genentech’s Giredestrant in ESR1-Mutated, ER-Positive Advanced Breast Cancer


Genentech, a member of the Roche Group (SIX: RO, ROG; OTCQX: RHHBY), announced  the U.S. Food and Drug Administration (FDA)  acceptance of  the company’s New Drug Application for giredestrant, an investigational oral therapy, in combination with everolimus for the treatment of adult patients with estrogen receptor (ER)-positive, human epidermal growth factor receptor 2-negative, ESR1-mutated locally advanced or metastatic breast cancer following recurrence or progression on a prior endocrine-based regimen. The FDA is expected to make a decision on the approval by December 18, 2026. Giredestrant plus everolimus could be the first and only oral selective estrogen receptor degrader (SERD) combination approved in the post-cyclin-dependent kinase (CDK)4/6 inhibitor setting.


Filing acceptance based on Phase III data showing giredestrant plus everolimus reduced the risk of disease progression or death by 44% and 62% in ITT and ESR1-mutated populations, respectively, versus standard-of-care endocrine therapy plus everolimus
Strength of evERA data demonstrate potential for giredestrant combination to help address resistance to standard-of-care therapies, and could be the first and only oral SERD combination approved in the post-CDK4/6 inhibitor setting
“The clinically meaningful benefit seen with giredestrant could enable an important new treatment option to help delay disease progression or death in people with advanced, ER-positive breast cancer,” said Levi Garraway, M.D., Ph.D., chief medical officer and head of Global Product Development. “This acceptance marks a first step towards establishing the giredestrant combination as a new standard of care in this population.”

The filing acceptance is based on the Phase III evERA Breast Cancer study results, which showed that giredestrant plus everolimus reduced the risk of disease progression or death by 44% and 62% in the intention-to-treat (ITT) and ESR1-mutated populations, respectively, compared with standard-of-care endocrine therapy plus everolimus. In the ESR1-mutated population, the median progression-free survival (PFS) was 9.99 months compared with 5.45 months in the giredestrant and comparator arm, respectively (stratified hazard ratio [HR]=0.38, 95% CI: 0.27-0.54, p-value=<0.0001). In the ITT population, the median PFS was 8.77 months compared with 5.49 months in the giredestrant and comparator arms, respectively (HR=0.56, 95% CI: 0.44-0.71, p-value=<0.0001).

Overall survival (OS) data were immature at the time of analysis, but a clear positive trend has been observed in the ITT (HR=0.69, 95% CI: 0.47-1.00, p-value=0.0473) and ESR1-mutated populations (HR=0.62, 95% CI: 0.38-1.02, p-value=0.0566). Follow-up for OS will continue to the next analysis. Adverse events for the giredestrant combination were manageable and consistent with the known safety profiles of the individual medicines. No unexpected safety findings were observed, including no photopsia.

Data from evERA are being used to support filing submissions to other global health authorities.

ER-positive breast cancer accounts for approximately 70% of breast cancer cases. Resistance to endocrine therapies, particularly in the post-CDK inhibitor setting, increases the risk of disease progression and is associated with poor outcomes. Oral combination therapies, such as giredestrant plus everolimus, could address this by targeting two different signaling pathways while helping to minimize the impact of treatment on people’s lives without the need for injections.

evERA was the first positive Phase III readout for giredestrant, followed by lidERA Breast Cancer in the early-stage setting. The scientific rationale for lidERA was supported by prior results in the neoadjuvant setting, including the coopERA trial showing that giredestrant was superior to an aromatase inhibitor in reducing malignant cell division (Ki67 levels). This growing body of evidence underscores the potential of giredestrant to become a new standard-of-care endocrine therapy across ER-positive early-stage and advanced breast cancer. In the coming weeks, Genentech will submit the giredestrant Phase III lidERA data in early-stage breast cancer to the FDA. The persevERA readout in first-line ER-positive breast cancer is expected in the first half of this year, which will provide further evidence for giredestrant in the ER-positive breast cancer treatment paradigm.

Our extensive giredestrant clinical development program spans multiple treatment settings and lines of therapy, reflecting our commitment to deliver innovative medicines to as many people with ER-positive breast cancer as possible.

https://en.wikipedia.org/wiki/Giredestrant


Wednesday, August 5, 2026

Nuvalent Announces Submission of New Drug Application to FDA for Neladalkib in TKI Pre-treated Advanced ALK-Positive NSCLC

Nuvalent, Inc. (Nasdaq: NUVL), a clinical-stage biopharmaceutical company focused on creating precisely targeted therapies for clinically proven kinase targets in cancer, today announced the submission to the U.S. Food and Drug Administration (FDA) of the Company's NDA for neladalkib, an investigational ALK-selective inhibitor, in TKI pre-treated advanced ALK-positive NSCLC.


New Drug Application (NDA) based on data in TKI pre-treated patients from the global ALKOVE-1 Phase 1/2 clinical trial
"The advancement of neladalkib from first clinical trial initiation to NDA submission in less than four years represents a remarkable pace in oncology drug development, underscoring the vigor and urgency our team brought to this program and our deep commitment to the ALK-positive NSCLC community," said Darlene Noci, A.L.M., Chief Development Officer at Nuvalent. "We would like to extend our sincere gratitude to the patients, families and investigators who have made this progress possible, and are committed to working closely with the FDA throughout the NDA review process toward our goal of bringing neladalkib to patients as quickly as possible."

The application is based on data in TKI pre-treated patients with advanced ALK-positive NSCLC treated with neladalkib in the global, registration-directed ALKOVE-1 Phase 1/2 clinical trial. In this population, neladalkib demonstrated encouraging overall activity, including intracranial responses, the ability to address key drivers of disease progression, and a generally well-tolerated safety profile consistent with its ALK-selective, TRK-sparing design. The company plans to share detailed results at a future medical meeting.

Neladalkib has received breakthrough therapy designation from the FDA for the treatment of patients with locally advanced or metastatic ALK-positive NSCLC who have been previously treated with 2 or more ALK tyrosine kinase inhibitors and orphan drug designation for ALK-positive NSCLC.

About Neladalkib
Neladalkib is an investigational, brain-penetrant, ALK-selective inhibitor created with the aim to overcome limitations observed with currently available ALK inhibitors. Neladalkib is designed to remain active in tumors that have developed resistance to first-, second-, and third-generation ALK inhibitors, including tumors with single or compound treatment-emergent ALK mutations such as G1202R. In addition, neladalkib is designed for central nervous system (CNS) penetrance to improve treatment options for patients with brain metastases, and to avoid inhibition of the structurally related tropomyosin receptor kinase (TRK) family. Together, these characteristics have the potential to avoid TRK-related CNS adverse events seen with dual TRK/ALK inhibitors and to drive deep, durable responses for patients across all lines of therapy. Neladalkib has received breakthrough therapy designation from the U.S. Food and Drug Administration (FDA) for the treatment of patients with locally advanced or metastatic ALK-positive non-small cell lung cancer (NSCLC) who have been previously treated with 2 or more ALK tyrosine kinase inhibitors and orphan drug designation for ALK-positive NSCLC.


https://go.drugbank.com/drugs/DB16688

Monday, August 3, 2026

Aldeyra Therapeutics Receives Complete Response Letter from the U.S. Food and Drug Administration for the Reproxalap New Drug Application for the Treatment of Signs and Symptoms of Dry Eye Disease

Aldeyra Therapeutics, Inc. (Nasdaq: ALDX) (Aldeyra), a biotechnology company devoted to discovering and developing innovative therapies designed to treat immune-mediated diseases, announced receipt of a Complete Response Letter (CRL) from the U.S. Food and Drug Administration (FDA) for the New Drug Application (NDA) of reproxalap, an investigational drug candidate, for the treatment of dry eye disease. The CRL stated that there is “a lack of substantial evidence consisting of adequate and well-controlled investigations … that the drug product will have the effect it purports or is represented to have under the conditions of use prescribed, recommended, or suggested in its proposed labeling” and that “the application has failed to demonstrate efficacy in adequate and well controlled studies in the treatment of signs and symptoms of dry eye disease.” The letter also stated that the “inconsistency of study results raises serious concerns about the reliability and meaningfulness of the positive findings” and that the “totality of evidence from the completed clinical trials does not support the effectiveness of the product.” Consistent with prior NDA reviews of reproxalap, no safety or manufacturing concerns were identified.


During the NDA review, label drafts were provided by the FDA in December 2025 and again in March 2026. Aldeyra does not believe that label negotiations were completed.

The FDA recommended that the reasons for failure in certain trials be explored, and that populations or certain conditions in which reproxalap may be effective be identified. The FDA did not recommend conducting additional trials or request submission of additional confirmatory evidence. As such, Aldeyra does not currently expect to pursue additional clinical trials, and intends to expeditiously request a Type A meeting to understand the actions needed for NDA approval. Per Prescription Drug User Fee Act (PDUFA) goals, the target Type A meeting date is within 30 days of receipt of the meeting request.

“To the thousands of American and Canadian patients who participated in our clinical trials and to the tens of millions of patients with dry eye disease worldwide, I want to assure you that we will work with urgency to support the FDA in enabling market access to what is, to our knowledge, the only drug with clinical activity within minutes of administration in patients with dry eye disease, a condition that is today treated with medications that require weeks or months of treatment to achieve even modest improvement,” stated Todd C. Brady, M.D., Ph.D., President and Chief Executive Officer of Aldeyra.

https://go.drugbank.com/drugs/DB16688

Saturday, August 1, 2026

Cogent Biosciences Announces Submission of New Drug Application for Bezuclastinib in Gastrointestinal Stromal Tumors (GIST)


Cogent Biosciences, Inc. (Nasdaq: COGT), a biotechnology company focused on developing precision therapies for genetically defined diseases, announced the completion of the submission of its New Drug Application (NDA) to the U.S. Food and Drug Administration (FDA) for bezuclastinib in patients with Gastrointestinal Stromal Tumors (GIST) who have received prior treatment with imatinib. Based on the positive results from the PEAK trial, the bezuclastinib NDA was submitted under the FDA’s Real-Time Oncology Review (RTOR) program, which is intended to enable a more streamlined review process. Bezuclastinib was also granted Breakthrough Therapy Designation as a treatment for GIST earlier this year.


Bezuclastinib NDA submitted under the FDA’s RTOR program based on positive results from Phase 3 PEAK trial; bezuclastinib previously granted Breakthrough Therapy Designation in GIST
Bezuclastinib combination demonstrated 16.5 month mPFS and 46% ORR in imatinib-resistant GIST patients, dramatically improving upon the current standard of care
“We are excited to complete our PEAK NDA submission which marks a significant step toward bringing a new therapy to patients with second-line GIST,” said Andrew Robbins, President and Chief Executive Officer. “Based on the strength of the PEAK data, we believe the bezuclastinib combination has the potential to meaningfully change the treatment landscape for these patients. We are grateful to the patients, investigators, and study teams who made this possible.”

Pivotal data from PEAK, a global, randomized Phase 3 clinical trial evaluating bezuclastinib in combination with sunitinib vs. sunitinib monotherapy in patients with GIST who have received prior treatment with imatinib, were reported in November 2025. As disclosed in the top-line results, the bezuclastinib combination demonstrated a substantial and highly statistically significant clinical benefit on the primary endpoint of progression free survival (PFS), reducing risk of disease progression or death compared to the current standard of care by 50% (hazard ratio of 0.50, 95% CI: 0.39 – 0.65). mPFS, as assessed by blinded independent central review, was 16.5 months for the bezuclastinib combination vs. 9.2 months for sunitinib monotherapy. Additionally, the bezuclastinib combination demonstrated an unprecedented ORR in imatinib-resistant patients, with 46% of patients treated with the bezuclastinib combination achieving an objective response compared to 26% of patients treated with sunitinib. The bezuclastinib combination was generally well tolerated, and no unique risks were observed with the novel combination when compared to the known safety profile of sunitinib. Data for overall survival remains immature.

At the time of data cutoff, based on the number of ongoing patients receiving treatment on the bezuclastinib combination arm, the estimated mean duration of treatment for the bezuclastinib combination is projected to exceed 19 months.

Cogent plans to present full results from the PEAK trial at a major medical meeting during the first half of 2026. Additionally, Cogent is on track this quarter to initiate a Phase 2 trial investigating the benefit of the bezuclastinib plus sunitinib combination for first-line GIST patients with exon 9 mutations who are naive to, or recently initiated treatment with, imatinib. The NDA submission with bezuclastinib in Advanced Systemic Mastocytosis (AdvSM) also remains on track for the first half of 2026.

Bezuclastinib - Expanded Access Program
Working with the FDA, Cogent has established active Expanded Access Programs (EAPs) for U.S. patients with GIST or SM who meet disease-specific criteria and could benefit from treatment with bezuclastinib or the combination of bezuclastinib and sunitinib. A growing number of sites now offer access to the bezuclastinib EAPs. For more information please visit: https://www.cogentbio.com/bezuclastinib-program-development/#our-expanded-access-policy

Inducement Grants Under Nasdaq Listing Rule 5635(c)(4)
Cogent also announced today that, on March 30, 2026, the Compensation Committee of Cogent’s Board of Directors, made up entirely of independent directors, approved the grants of “inducement” equity awards to six new employees under the company’s 2020 Inducement Plan with a grant date of March 30, 2026. The awards were approved in accordance with Listing Rule 5635(c)(4) of the corporate governance rules of the Nasdaq Stock Market. The employees received, in the aggregate, (i) nonqualified options to purchase 21,100 shares of Cogent common stock and (ii) 15,700 restricted stock units (RSUs). Each option has a 10-year term, an exercise price equal to the closing price of Cogent’s common stock on the grant date, and a 4-year vesting schedule with 25% vesting on the 1-year anniversary of the grant date and the remainder vesting in equal monthly installments over the subsequent 36 months, provided such employee remains employed through each such vesting date. The RSUs vest annually in equal installments over 4 years from the grant date, provided such employee remains employed through each such vesting date.

https://en.wikipedia.org/wiki/Bezuclastinib

Tuesday, June 30, 2026

Praxis Precision Medicines Announces FDA Acceptance and Priority Review of New Drug Application for Relutrigine in Patients with SCN2A and SCN8A DEEs

Praxis Precision Medicines, Inc. (NASDAQ: PRAX), a fully integrated, leading central nervous system (CNS) precision neuroscience biopharmaceutical company, today announced that the U.S. Food and Drug Administration (FDA) has accepted for priority review its New Drug Application (NDA) for relutrigine, for the treatment of SCN2A and SCN8A developmental and epileptic encephalopathies (DEEs). The FDA has set a target action date under the Prescription Drug User Fee Act (PDUFA) of September 27, 2026.


  • FDA assigned PDUFA target action date of September 27, 2026

“Our first FDA acceptance of an NDA submission marks a significant milestone in our evolution to a commercial-stage company and an important step toward delivering innovative, precision neuroscience therapies to patients in need. SCN2A/8A DEEs have no currently approved targeted therapies and relutrigine, if approved, would be the first disease-modifying therapy for children suffering from these devastating and fatal conditions. We look forward to working closely with the FDA during the review process while continuing to advance our launch preparations,” said Marcio Souza, president and chief executive officer.

Relutrigine for treatment of SCN2A/8A DEEs
The NDA is supported by positive results from the EMBOLD study, which was stopped early for efficacy following a successful interim analysis and recommendation from the Data Monitoring Committee. Relutrigine has an Orphan Drug Designation, as well as a Rare Pediatric Disease Designation and a Breakthrough Therapy Designation. If granted approval, relutrigine will be the first FDA-approved therapy for SCN2A/8A DEE as well as be eligible for a Pediatric Review Voucher.

Relutrigine is also being investigated in broad DEEs through the EMERALD trial, which is expected to be completed by the end of 2026.

About Relutrigine
Relutrigine is a first-in-class small molecule in development for the treatment of developmental and epileptic encephalopathies (DEEs) as a preferential inhibitor of persistent sodium current, shown to be a key driver of seizure symptoms in severe DEEs. Relutrigine’s mechanism of precision sodium channel (NaV) modulation is consistent with superior selectivity for disease-state NaV channel hyperexcitability. In vivo studies of relutrigine have demonstrated dose-dependent inhibition of seizures up to complete control of seizure activity in SCN2A, SCN8A and other DEE mouse models. Relutrigine has been generally well-tolerated in three Phase 1 studies and has demonstrated biomarker changes indicative of NaV channel modulation. Data from cohort 1 of the Phase 2 EMBOLD study demonstrated a well-tolerated, robust, short- and long-term improvement in motor seizures in a heavily pre-treated population, alongside maintained seizure freedom in some patients with SCN2A- and SCN8A-DEE. Relutrigine has received Orphan Drug Designation (ODD) and Rare Pediatric Disease Designation from the FDA for the treatment of SCN2A-DEE, SCN8A-DEE and Dravet syndrome; as well as Breakthrough Therapy Designation (BTD), and ODD from the European Medicines Agency for the treatment of SCN2A-DEE and SCN8A-DEE.


Monday, June 29, 2026

Elevar Therapeutics Announces FDA Acceptance for Review of New Drug Application for Lirafugratinib as Second-line Cholangiocarcinoma Treatment

Elevar Therapeutics, Inc., a majority-owned subsidiary of HLB Co., Ltd. and a fully integrated biopharmaceutical company dedicated to elevating treatment experiences and outcomes for patients who have limited or inadequate therapeutic options, today announced that the U.S. Food and Drug Administration (FDA) has completed its filing review of the New Drug Application (NDA) for lirafugratinib, an investigational therapy, for the treatment of patients with cholangiocarcinoma (CCA) with FGFR2 fusions or rearrangements who have received prior therapy. The FDA determined that the NDA is sufficiently complete to permit a substantive review and granted Priority Review, with a Prescription Drug User Fee Act (PDUFA) target action date of September 27, 2026.


  • With Priority Review designation, FDA set a Sept. 27 PDUFA date for approval decision
  • Lirafugratinib achieved a 46.5% ORR, in CCA patients with FGFR2 fusion and rearrangement

Priority Review designations are given to applications for medicines that, if approved, would lead to “significant improvements in the safety or effectiveness of the treatment” of a serious condition, according to the FDA. The Priority Review designation of the lirafugratinib NDA was supported by positive clinical data from the Phase 1/2 ReFocus trial (NCT04526106), which demonstrated a confirmed objective response rate (ORR) of 46.5% in the patients with the proposed indication. Its safety profile in the clinical data has been shown to be predictable and manageable through dose adjustments.

“Lirafugratinib has established a compelling clinical profile that differentiates it from existing treatment options,” said Dong-Gun Kim, chief executive officer of Elevar. “We are very pleased with the FDA’s priority review designation and focused on advancing the review process efficiently to bring this therapy to patients as quickly as possible.”

CCA, also known as bile duct cancer, is rare, with about 8,000 people in the U.S. diagnosed each year, according to the American Cancer Society.

Elevar Therapeutics continues to evaluate lirafugratinib in ongoing clinical development programs, including studies in other FGFR2-altered solid tumors. Any future indications will be subject to regulatory review and approval.

For more information about lirafugratinib, visit ElevarTX.com.

About Lirafugratinib

Lirafugratinib (RLY-4008) is a potent, selective, and oral small molecule inhibitor of FGFR2, a receptor tyrosine kinase that is frequently altered in certain cancers. FGFR2 is one of four members of the FGFR family, a set of closely related proteins with highly similar protein sequences and properties. Preclinically, lirafugratinib demonstrated FGFR2-dependent killing in cancer cell lines and induced regression in in vivo models with minimal inhibition of other targets, including other members of the FGFR family. In addition, lirafugratinib demonstrated strong activity against known clinical on-target resistance mutations in vitro and in vivo preclinical models. Lirafugratinib is currently being evaluated in a clinical trial to enroll additional patients with previously treated, advanced or metastatic solid tumors other than CCA harboring FGFR2 fusion or rearrangement, who have not been treated with prior FGFR inhibitors.


Saturday, June 6, 2026

U.S. Food and Drug Administration Accepts Bristol Myers Squibb's New Drug Application for Iberdomide in Patients with Relapsed or Refractory Multiple Myeloma


Bristol Myers Squibb (NYSE: BMY) today announced that the U.S. Food and Drug Administration (FDA) has accepted a New Drug Application (NDA) for iberdomide combined with standard treatment (daratumumab + dexamethasone - IberDd) in patients with relapsed or refractory multiple myeloma (RRMM). Iberdomide is part of an investigational, new class of medicines called cereblon E3 ligase modulator (CELMoD) agents. The FDA has granted a Prescription Drug User Fee Act (PDUFA) date of August 17, 2026 for this indication.


  • Iberdomide has the potential to be the first approved CELMoD agent
  • The U.S. FDA has granted Breakthrough Therapy Designation and Priority Review for this indication and assigned a target action date of August 17, 2026

“The FDA’s acceptance of this application is a testament to the potential of iberdomide, in combination with anti-CD38 monoclonal antibodies, as a novel, potent, oral treatment option, with a manageable safety profile, for patients with multiple myeloma,” said Cristian Massacesi, executive vice president and chief medical officer, Bristol Myers Squibb. “Furthermore, our filing for iberdomide based on the MRD endpoint, underscores our commitment to pioneering new ways of advancing life-saving therapies for patients living with cancer.”

The filing was based on results from a planned analysis of MRD negativity rates in the Phase 3 EXCALIBER-RRMM study evaluating iberdomide as a treatment for RRMM patients. The EXCALIBER-RRMM trial is ongoing and patients continue to be evaluated for progression-free survival (PFS).

The FDA also granted Breakthrough Therapy designation for iberdomide based on these data.

This review is being conducted under the FDA’s Project Orbis initiative, which enables concurrent review by the health authorities in several other countries.

https://en.wikipedia.org/wiki/Iberdomide


Friday, June 5, 2026

Deciphera Pharmaceuticals Announces U.S. Food and Drug Administration Acceptance for Filing of New Drug Application for Tirabrutinib in Patients with Relapsed or Refractory PCNSL

Deciphera Pharmaceuticals, a member of Ono Pharmaceutical Co., Ltd. (Headquarters: Osaka, Japan; President and COO: Toichi Takino; “Ono”), today announced that the U.S. Food and Drug Administration (FDA) has accepted for filing the New Drug Application (NDA) under the accelerated approval pathway for tirabrutinib, a highly selective irreversible, second generation Bruton tyrosine kinase inhibitor, for the treatment of relapsed or refractory primary central nervous system lymphoma (R/R PCNSL). The FDA has set an action date of December 18, 2026, under the Prescription Drug User Fee Act (PDUFA).


Regulatory submission is based on positive results from the Phase 2 PROSPECT Study
FDA sets PDUFA Date of December 18, 2026
“R/R PCNSL is a rare and aggressive form of non-Hodgkin lymphoma with particularly poor clinical outcomes. Patients often experience difficulty and delay in diagnosis, and once they are diagnosed, there is a high unmet need for a treatment with a favorable safety profile,” said Matthew L. Sherman, M.D., Chief Medical Officer of Deciphera. “The FDA’s acceptance of tirabrutinib’s NDA for filing is an exciting milestone as it brings us one step closer to our goal of providing patients with R/R PCNSL an important new treatment option.”

“We are very pleased that the NDA for tirabrutinib has been accepted for filing,” said Toichi Takino, President and COO of Ono Pharmaceutical Co., Ltd. “This is an important milestone on the way to expanding our commercial pipeline and achieving our goal of becoming a global specialty pharma. Tirabrutinib’s potential to address unmet patient needs embodies our corporate philosophy and we will continue to focus on developing and delivering innovative medicines to benefit patients worldwide.”

The NDA is supported by the positive results from the Phase 2 PROSPECT study, presented at the 2025 American Society for Clinical Oncology (ASCO) Annual Meeting, in which tirabrutinib demonstrated an overall response rate of 67%, a complete response rate of 44%, and a manageable safety profile. If approved, tirabrutinib will be the first BTK inhibitor therapy commercially available in the U.S. for the treatment of patients with R/R PCNSL, and the third commercial therapy for Ono group available in the U.S. The study is currently recruiting patients with R/R PCNSL in a global Phase 3 randomized trial, which will serve as a confirmatory study for this indication (ClinicalTrials.gov NCT07104032.)


https://en.wikipedia.org/wiki/Tirabrutinib

Saturday, April 11, 2026

Arvinas Announces FDA Acceptance of the New Drug Application for Vepdegestrant for the Treatment of ESR1m, ER+/HER2- Advanced Breast Cancer

Arvinas, Inc. (Nasdaq: ARVN), today with its partner Pfizer Inc. (NYSE: PFE), announced that the U.S. Food and Drug Administration (FDA) has accepted the New Drug Application (NDA) for vepdegestrant for the treatment of patients with estrogen receptor-positive (ER+)/human epidermal growth factor receptor 2-negative (HER2-), ESR1-mutated advanced or metastatic breast cancer who have previously received endocrine-based therapy. The FDA has assigned a Prescription Drug User Fee Act (PDUFA) action date of June 5, 2026.


“Patients often face limited treatment options after first-line treatment and vepdegestrant demonstrated improved progression-free survival in patients with ESR1-mutated ER+/HER2- advanced breast cancer,” said John Houston, Ph.D., Chairperson, Chief Executive Officer, and President at Arvinas. “With the efficacy and favorable tolerability seen in VERITAC-2, we believe vepdegestrant, if approved, has potential to be a best-in-class treatment option for patients in the second-line ESR1-mutant setting. We look forward to working alongside Pfizer and with the FDA to pursue vepdegestrant’s approval and to ensure this important treatment option is made available to patients as rapidly as possible.”

Vepdegestrant, an investigational oral PROTAC ER degrader, is being jointly developed by Arvinas and Pfizer. The NDA submission was based on data from VERITAC-2 (NCT05654623), a global, randomized Phase 3 clinical trial evaluating vepdegestrant versus fulvestrant. These data were recently presented at the American Society of Clinical Oncology (ASCO) 2025 Annual Meeting and were simultaneously published in The New England Journal of Medicine.

About Vepdegestrant
Vepdegestrant is an investigational, orally bioavailable PROteolysis TArgeting Chimera (PROTAC) estrogen receptor degrader. Vepdegestrant is being developed as a potential monotherapy for ER+/HER2- advanced or metastatic breast cancer with estrogen receptor 1 (ESR1) mutations in the second line-plus setting.

In July 2021, Arvinas announced a global collaboration with Pfizer for the co-development and co-commercialization of vepdegestrant; Arvinas and Pfizer will share worldwide development costs, commercialization expenses, and profits.

The U.S. Food and Drug Administration (FDA) has accepted the New Drug Application (NDA) for vepdegestrant for its use as a monotherapy in the treatment of adults with estrogen receptor–positive (ER+), human epidermal growth factor receptor 2–negative (HER2-), ESR1-mutated advanced or metastatic breast cancer previously treated with endocrine-based therapy. Vepdegestrant has also been granted Fast Track designation by the FDA, underscoring the significant unmet need in this patient population and the potential for vepdegestrant to offer a meaningful new treatment option.

About the VERITAC-2 Clinical Trial
The Phase 3 VERITAC-2 clinical trial (NCT05654623) is a global, randomized trial evaluating the efficacy and safety of vepdegestrant (ARV-471) as a monotherapy compared to fulvestrant in patients with ER+/HER2- advanced or metastatic breast cancer previously treated with a CDK4/6 inhibitor plus endocrine therapy. The trial enrolled 624 patients, 270 of whom had ESR1m positive disease, at 213 sites in 25 countries.

Patients were randomized 1:1 to receive either vepdegestrant once daily, orally on a 28-day continuous dosing schedule, or fulvestrant, administered intramuscularly on Days 1 and 15 of Cycle 1 and then on Day 1 of each 28-day cycle starting from Day 1 of Cycle 2. In the trial, 43% of patients (n=270) had ESR1 mutations detected. The primary endpoint was progression-free survival (PFS) in the ESR1-mutation and intent-to-treat populations as determined by blinded independent central review. Overall survival is the key secondary endpoint.

https://en.wikipedia.org/wiki/Vepdegestrant

Wednesday, April 1, 2026

Achieve Life Sciences Announces FDA Acceptance of Cytisinicline New Drug Application for Treatment of Nicotine Dependence for Smoking Cessation

Achieve Life Sciences, Inc. (Nasdaq: ACHV), a late-stage specialty pharmaceutical company focused on the global development and commercialization of cytisinicline for treatment of nicotine dependence, today announced that the U.S. Food and Drug Administration (FDA) has accepted the cytisinicline New Drug Application (NDA) for a new treatment for smoking cessation in adults. The FDA has assigned a Prescription Drug User Fee Act (PDUFA) targeted action date of June 20, 2026.


“The FDA’s acceptance of our NDA validates the totality of our product development program,” said Dr. Cindy Jacobs, President and Chief Medical Officer of Achieve Life Sciences. “Our application is supported by a decade of rigorous research and comprehensive data from thousands of participants. We've built compelling scientific and clinical results as a foundation that, if approved, positions cytisinicline to potentially address a significant medical need. We’re eager to engage constructively with the FDA as we progress through the NDA review process.”

The company’s NDA details a comprehensive clinical development program, with more than 2,000 clinical trial participants contributing to the body of evidence. The ORCA-2 and ORCA-3 Phase 3 trials showed cytisinicline administered for either 6 or 12 weeks, alongside standard behavioral support, demonstrated significantly greater smoking abstinence rates by the end of treatment and in long-term abstinence through week 24 as compared to placebo. In addition, safety data will include over 400 participants who have received at least six months of cumulative cytisinicline exposure and over 200 participants receiving at least 1 year of cumulative cytisinicline exposure, with no new safety concerns as reported by the Data Safety Monitoring Committee.

“Smoking is the leading cause of preventable death and disease, claiming the lives of nearly half a million Americans each year, and costing the American economy more than an estimated $600 billion a year,” said Rick Stewart, Chief Executive Officer of Achieve Life Sciences. “The FDA's acceptance of our NDA filing underscores nicotine dependence as an important public health need that demands action. We're energized by this milestone and remain laser-focused on our mission, working toward commercial readiness for the second half of 2026, pending FDA approval. Each year, an estimated 15 million Americans attempt to quit smoking, and we're committed to providing those who are ready to quit a new tool so they can break free from nicotine dependence.”

https://en.wikipedia.org/wiki/Cytisine

Thursday, February 19, 2026

Sentynl Therapeutics Updates, Complete Response Letter (CRL) relating to its New Drug Application (NDA) for On Its NDA for CUTX-101

Sentynl Therapeutics, Inc. (“Sentynl”), a U.S.-based biopharmaceutical company wholly-owned by Zydus Lifesciences, Ltd. (“Zydus”), announced today that the U.S. Food and Drug Administration (USFDA) has issued a Complete Response Letter (CRL) relating to its New Drug Application (NDA) for copper histidinate (CUTX-101), intended to treat Menkes disease in pediatric patients.

The USFDA provided findings within the CRL that Sentynl and Zydus will need to address to clarify the path forward. Specifically, the USFDA mentioned a CGMP inspection of the facility where CUTX-101 is manufactured. Zydus recently provided responses to USFDA’s September 2025 re-inspection demonstrating the facility’s CGMP compliance and is awaiting USFDA’s Establishment Inspection Report (EIR). Sentynl will request a meeting with USFDA to discuss the CRL and resubmission of the CUTX-101 NDA. The CRL did not cite any other approvability concerns, nor did it identify any deficiencies in CUTX-101’s efficacy and safety data.

“We recognize the USFDA’s decision and remain dedicated to working with the Agency to clarify next steps. Our commitment to patients is unchanged. We believe in the promise of our therapy and are prepared to address the feedback and pursue resubmission promptly,” said Matt Heck, CEO, Sentynl.

Menkes disease is a rare X-linked recessive pediatric genetic disease that impacts an estimated 1 in 34,810 to as high as 1 in 8,664 live male births. Patients with Menkes disease are born with the inability to absorb dietary copper and subsequently have impaired copper transport across the blood-brain barrier. CUTX-101 is a subcutaneous injectable formulation of copper histidinate that restores copper homeostasis and maintains copper levels in patients with Menkes disease.

The CUTX-101 NDA was initially granted Priority Review by the FDA and is supported by positive topline clinical efficacy results for CUTX-101, demonstrating significant improvement in overall survival for Menkes disease subjects who received early treatment with CUTX-101.


Sentynl Therapeutics Updates On Its NDA for CUTX-101

Tuesday, February 3, 2026

FDA Issues Complete Response Letter for Biohaven's Vyglxia (troriluzole) New Drug Application for Spinocerebellar Ataxia


Biohaven Ltd. (NYSE: BHVN) ("Biohaven"), a global clinical-stage biopharmaceutical company focused on the discovery, development and commercialization of life-changing therapies to treat a broad range of rare and common diseases,  announced  it has received a Complete Response Letter (CRL) from the U.S. Food and Drug Administration (FDA) for the New Drug Application (NDA) seeking approval of Vyglxia (troriluzole) for the treatment of spinocerebellar ataxia (SCA).


Vlad Coric, M.D., Chairman and Chief Executive Officer of Biohaven said, "We are extremely disappointed on behalf of patients by this action from the Office of Neuroscience at FDA. Beyond substantial evidence of safety and efficacy, patients with rare diseases also deserve an efficient, fair and flexible regulatory process that aligns with the urgency of their high unmet medical needs. Such an approach has been mandated by Congress to empower the FDA with maximum regulatory flexibility for rare disease. As a company, we are committed to advancing innovative treatments and remain dedicated to SCA patients despite all the challenges associated with pursuing therapies for rare diseases. Real-world evidence is an important research approach to assessing and delivering new therapies for complex rare diseases but, despite FDA policy initiatives supporting such tools, the front-line review divisions are not yet embracing FDA policy for the use of real-world evidence or the application of regulatory flexibility for rare disease."

Jeremy Schmahmann, M.D., Professor of Neurology at Harvard Medical School and Founding Director of the Ataxia Unit at Massachusetts General Hospital (MGH), added, "Patients with SCA and clinicians who treat them deserve to be heard on this important NDA filing. There is too much at stake for patients. The FDA decision not to listen to disease experts and respect the patient perspective before taking action represents a misstep in the due process, and a failure to deploy regulatory flexibility to evaluate benefit:risk of a medication that has proven to be safe and effective for this rare, debilitating neurodegenerative disease that has no current treatment."

Dr. Coric added, "The development of Vyglxia® (troriluzole) by Biohaven embodies a strong scientific process and deep commitment that is critical to bringing safe and effective treatments to patients with rare diseases like SCA. Our efforts over eight years, included developing the f-SARA scale in collaboration with the FDA and a real-world evidence study in SCA that showed Vyglxia achieved highly consistent, sustained, robust and clinically meaningful treatment effects with a safe, once-daily oral pill that slowed disease progression by 50-70%. The NDA also included data showing Vyglxia reduced the risk of falls and delayed time to becoming wheelchair bound. The leading SCA experts in the United States directly communicated their support of the troriluzole data to the FDA but unfortunately the Office of Neuroscience's inability to collaboratively engage with Biohaven, the patient community and leading experts leave us with concerns about the lack of regulatory flexibility that is being applied for rare, life-threatening conditions. There are a number of common sense solutions and regulatory tools that the Office of Neuroscience could have applied including a fair hearing of the drug's efficacy and safety risks at an Advisory Committee of experts and patients, post-marketing studies, labelling limitations or an accelerated approval pathway. Patients are waiting and the certainty of disease progression for SCA patients far outweighs any residual uncertainty regarding potential design bias or interpretation of study data, especially when the primary outcome measure was achieved in a study protocol and statistical analysis plan that was reviewed by the FDA prior to data analysis. SCA patients deserved approval of Vyglxia and certainly a more balanced interpretation of benefit:risks."

Biohaven remains committed to working with the FDA to find a path forward for its NDA for Vyglxia and plans to meet with the FDA to discuss potential next steps.

Prioritizing Clinical-Stage, Innovative Assets

Biohaven will prioritize resources to focus all its R&D resources on other key programs from its diversified portfolio. Consistent with Biohaven's enduring commitment as a patient-first drug developer, the company's pipeline is focused on a range of disease indications which have limited or no treatment options and are long overdue for therapeutic innovation.

Bruce Car Ph.D., Chief Scientific Officer at Biohaven, commented, "As drug developers we expect setbacks and our diversified portfolio affords us the opportunity to pivot to other key programs. We remain as resilient as ever in following science in order to make a difference in the lives of people with debilitating diseases. Much important work remains, and we are energized and focused on achieving the critical milestones that lie ahead, mindful that days matter and patients are waiting."

Biohaven is initiating strategic portfolio and cost optimization across multiple programs and will focus forward-looking spend on restructuring of business priorities to achieve an approximately 60% reduction in annual direct R&D spend (which excludes personnel and SBC). This may include pausing or delaying non-priority programs to maintain its cash runway to focus on the priority clinical-stage programs over the next year.

https://en.wikipedia.org/wiki/Troriluzole


https://www.drugs.com/nda/vyglxia_251105.html

Friday, August 16, 2024

Zevra Therapeutics Announces Resubmission of Arimoclomol New Drug Application to the U.S. Food and Drug Administration


Zevra Therapeutics, Inc. (NasdaqGS: ZVRA) a rare disease therapeutics company, today announced it resubmitted its New Drug Application (NDA) for arimoclomol, an investigational therapeutic candidate for the treatment of Niemann-Pick disease type C (NPC) to the U.S. Food and Drug Administration (FDA) on December 22, 2023. Based on standard NDA resubmission review timelines, an acknowledgment letter from the FDA that the resubmission is complete and setting the PDUFA date is expected within 30 days. Zevra expects the NDA to be classified as a Class II submission which would be subject to a review period by the FDA within six months from the date of submission.

“The Zevra team has worked diligently to deliver a high quality and thorough resubmission of the NDA for arimoclomol following multiple interactions with the FDA and after incorporating direction from the agency,” said Neil McFarlane, President and Chief Executive Officer of Zevra. “We continue to accelerate our launch preparations in anticipation of FDA approval, and believe we are one step closer to getting arimoclomol into the hands of patients who are seeking a treatment.”

“Zevra has engaged with the advocacy community, elevating the patient voice throughout arimoclomol’s development process,” said Daniel Gallo, Ph.D., Zevra’s Senior Vice President of Medical Affairs and Advocacy. “The advocacy community’s input has been instrumental in building awareness of the need for approved treatments that address the unmet needs of individuals and their caregivers living with this debilitating condition.”

Zevra believes it has addressed the issues previously raised by the FDA in the complete response letter by providing additional evidence to support the use of the Niemann-Pick type C Clinical Severity Scale (NPCCSS) and, conducting additional studies used to support the potential mechanism of action. Additionally, new data included in the resubmission comes from multiple non-clinical studies, natural history comparisons, real-world data generated from the ongoing early access programs in the United States and the European Union, as well as data from the four-year open-label extension of the Phase 2/3 clinical trial (NCT02612129). Results from this open-label trial suggest that arimoclomol reduces the long-term progression of NPC.

Arimoclomol has been evaluated in a total of 21 studies across a range of Phase 1, 2 or 3 clinical trials evaluating its safety and efficacy across more than 600 subjects in NPC, other disease or healthy subjects. The primary efficacy trial evaluating arimoclomol for the treatment of NPC was a Phase 2/3 double-blind, placebo-controlled trial (CT-ORZY-NPC-002) of arimoclomol in 50 patients with NPC.


https://en.wikipedia.org/wiki/Arimoclomol

Thursday, August 15, 2024

Zealand Pharma Submits New Drug Application to the US FDA for Glepaglutide in Short Bowel Syndrome


Zealand Pharma A/S  announced the submission of a New Drug Application (NDA) to the US Food and Drug Administration (FDA) for glepaglutide, a long-acting GLP-2 analog, for the treatment of adult patients with short bowel syndrome (SBS) dependent on parenteral support.


“Short bowel syndrome with intestinal failure is a complex, chronic and severe condition in which individuals are dependent on receiving fluids and nutrition parenterally. While life-sustaining, parenteral support poses significant restrictions on daily life and carries a risk of serious and life-threatening complications. More effective and convenient treatments to further reduce parenteral support are needed, with the ultimate goal of discontinuing parenteral support and achieving enteral autonomy,” said David Kendall, MD, Chief Medical Officer of Zealand Pharma. “We believe glepaglutide, once approved, can reduce both the burden of parenteral support and of daily dosing of existing GLP-2 treatment for people living with SBS and intestinal failure, and we are pleased to submit this treatment for regulatory review and potential approval in the US.”

https://en.wikipedia.org/wiki/Sotorasib

Wednesday, August 14, 2024

Journey Medical Corporation Submits New Drug Application to FDA for DFD-29 to Treat Rosacea


Journey Medical Corporation,  announced that the Company has submitted,  a New Drug Application (“NDA”) to the FDA seeking approval for DFD-29 (Minocycline Hydrochloride Modified Release Capsules, 40 mg) for the treatment of inflammatory lesions and erythema of rosacea in adults. DFD-29 is being developed in collaboration with Dr. Reddy’s Laboratories Ltd.


“This NDA submission is a significant milestone for Journey Medical and we look forward to collaborating with the FDA during its review to bring DFD-29, a potentially differentiated, best-in-class oral rosacea treatment, one step closer to patients. Based on the data seen in our pivotal trials, DFD-29 could fundamentally improve the treatment paradigm for patients suffering from both inflammatory lesions and erythema (redness) from rosacea,” said Claude Maraoui, Co-Founder, President and Chief Executive Officer of Journey Medical.

The NDA submission is supported by positive data from Journey Medical’s two DFD-29 Phase 3 clinical trials for the treatment of rosacea. The Phase 3 clinical trials achieved all co-primary and secondary endpoints, and subjects completed the 16-week treatment with no significant safety issues. DFD-29 demonstrated statistical superiority over both the current standard-of-care treatment, Oracea® 40 mg capsules, and placebo for Investigator’s Global Assessment treatment success as well as the reduction in the total inflammatory lesion count in both studies. On a secondary endpoint related to erythema (redness) assessment, DFD-29 showed statistically significant reduction in Clinician’s Erythema Assessment (CEA) compared to placebo in both clinical trials.

Rosacea, which affects approximately 16 million people in the United States according to the National Rosacea Society, is a long-term, inflammatory skin condition that causes small, red, pus-filled bumps, redness and visible blood vessels in the face. DFD-29 has the potential to become the only oral, systemic therapy to address both inflammatory lesions and erythema (redness) from rosacea, as demonstrated in the clinical trials.

Ref : https://en.wikipedia.org/wiki/Minocycline

Friday, August 9, 2024

Amneal Announces Complete Response Resubmission for IPX203 New Drug Application

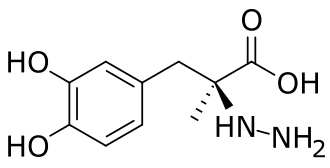
Amneal Pharmaceuticals, Inc. (NASDAQ: AMRX) (“Amneal” or the “Company”) today announced that it has provided a Complete Response resubmission to the U.S. Food and Drug Administration (FDA) for IPX203, a novel, oral formulation of carbidopa/levodopa (CD/LD) extended-release capsules for the treatment of Parkinson’s disease (PD).

Carbidopa
levodopa 


The original NDA for IPX203 resulted in a Complete Response Letter (CRL) from FDA. The resubmission package included data from a healthy volunteer study which was conducted in the fourth quarter of 2023. The FDA did not request any other studies.

“We are pleased to provide our complete response resubmission for IPX203 as we look to expand our Parkinson’s franchise,” said Chirag and Chintu Patel, Co-Chief Executive Officers at Amneal. “We look forward to launching this much-needed treatment in the second half of 2024, subject to FDA approval.”

About IPX203 

IPX203 is a novel, oral formulation of CD/LD extended-release capsules designed for the treatment of Parkinson’s disease. IPX203 contains immediate-release granules and extended-release coated beads. The IR granules consist of CD and LD, with a disintegrant polymer to allow for rapid dissolution. The ER beads consist of LD, coated with a sustained release polymer to allow for slow release of the drug a mucoadhesive polymer to keep the granules adhered to the area of absorption longer, and an enteric coating to prevent the granules from disintegrating prematurely in the stomach. This formulation is distinct from RYTARY® (carbidopa/levodopa) extended-release capsules, Amneal’s extended-release CD/LD treatment for PD approved by the U.S. FDA in 2015.

About Parkinson’s Disease
Parkinson’s disease has become the fastest growing neurological disorder worldwide, with approximately 1 million patients diagnosed in the U.S. It is a progressive disorder of the central nervous system (CNS) that affects dopamine-producing neurons in the brain that affect movement.

PD is characterized by slowness of movement, stiffness, resting tremor and impaired balance.  While PD is not considered a fatal disease, it is associated with significant morbidity and disability.  The average age at diagnosis for patients with PD is 60; as people live longer, the number of patients living with PD is predicted to grow significantly over the coming decades. 


https://en.wikipedia.org/wiki/Carbidopa
https://en.wikipedia.org/wiki/L-DOPA

Amneal Announces Complete Response Resubmission for IPX203 New Drug Application