Tuesday, September 8, 2026
FDA Approves Veppanu (vepdegestrant) for the Treatment of ESR1m, ER+/HER2- Advanced Breast Cancer
Monday, August 31, 2026
FDA Approves Idvynso (doravirine and islatravir) for the Treatment of HIV-1 Infection in Adults
In continuation of my update on Doravirine
Merck (NYSE: MRK), known as MSD outside of the United States and Canada, announced the U.S. Food and Drug Administration (FDA) approval of Idvynso, a new, two-drug single-tablet regimen of 100 mg doravirine and 0.25 mg islatravir, for the treatment of HIV-1 infection in adults to replace the current antiretroviral regimen in those who are virologically suppressed (HIV-1 RNA less than 50 copies per mL) on a stable antiretroviral regimen with no history of virologic treatment failure and no known substitutions associated with resistance to doravirine. Idvynso is contraindicated when co-administered with drugs that are strong cytochrome P450 (CYP)3A enzyme inducers and lamivudine (3TC) or emtricitabine (FTC). Co-administration with these drugs may decrease the effectiveness of Idvynso. See additional selected safety information on the following pages. Idvynso (pronounced ihd-VIHN-soh) will be available in pharmacies after May 11.
Islatravir
- Idvynso is approved for adults with virologically suppressed HIV-1 with no history of virologic treatment failure and no known substitutions associated with resistance to doravirine
- Idvynso is the first and only non-INSTI, tenofovir-free, once-daily, complete two-drug regimen to demonstrate non-inferior efficacy in a head-to-head Phase 3 trial versus three-drug regimen Biktarvyi (BIC/FTC/TAF)
“Advances in HIV treatment mean more people living with HIV are living longer — a remarkable achievement,” said Carl Baloney, Jr., president and chief executive officer of AIDS United. “People aging with HIV face additional health challenges, including managing multiple chronic conditions and medications at the same time. It is essential that management of HIV considers these factors in addition to virologic suppression when choosing an HIV treatment regimen.”
“Idvynso combines islatravir, a next-generation NRTI with multiple mechanisms of action, including translocation inhibition, with doravirine, an NNRTI with an established efficacy and safety profile. As the only two-drug, non-INSTI, tenofovir-free regimen, Idvynso expands therapeutic diversity beyond the currently available oral treatment options,” said Dr. Eliav Barr, senior vice president and chief medical officer, Merck Research Laboratories. “As the health needs of adults living with HIV change over time, Idvynso gives clinicians a new choice for HIV treatment. This approval marks an important new chapter in Merck’s long-standing commitment to research and discovery for people living with HIV.”
Idvynso is a complete regimen; co-administration with other antiretroviral medications for treatment of HIV-1 infection is not recommended. Severe skin reactions, including Stevens-Johnson syndrome/toxic epidermal necrolysis, have been reported with doravirine-containing regimens. Drug Rash with Eosinophilia and Systemic Symptoms was reported with Idvynso. Concomitant use of Idvynso and certain other drugs may result in known or potentially significant drug interactions, some of which may lead to loss of therapeutic effect of Idvynso and possible development of resistance, or possible clinically significant adverse reactions from greater exposures of a component of Idvynso. See additional selected safety information on the following pages.
“Idvynso is the first non-INSTI, tenofovir-free, two-drug regimen to demonstrate non-inferior efficacy to standard oral antiretroviral regimens, including Biktarvy. This makes Idvynso a potential alternative for people with virologically suppressed HIV who may need to switch their treatment,” said Dr. Amy Colson, director of research at Community Resource Initiative, Boston, Massachusetts.
Phase 3 studies supporting approval of Idvynso
The efficacy and safety of Idvynso is supported by Week 48 data from two randomized, active-controlled, non-inferiority trials [Trial 052 (NCT05630755) and Trial 051 (NCT05631093)] in virologically-suppressed (HIV-1 RNA less than 50 copies per mL) adults living with HIV. Participants must have been stably suppressed on their baseline regimen for at least 3 months prior to trial entry and had no history of treatment failure. Across the two trials, a total of 708 participants received once-daily Idvynso; of these, 81 (11%) participants were aged 65 years and older, including 10 (1%) aged 75 years and older.
In the double-blind Trial 052, participants were switched from Biktarvy [bictegravir/emtricitabine/tenofovir alafenamide (BIC/FTC/TAF)] to Idvynso. A total of 513 participants were randomized (2:1) and were switched to once-daily Idvynso (n=342) or remained on BIC/FTC/TAF (n=171). At baseline, participants had a mean age of 48 years (range: 19 to 77), 21% of participants were female, 61% were White, 31% were Black/African American, and 6% were Asian. A total of 23% identified as Hispanic/Latino.
In the open-label Trial 051, participants were switched from an oral ART (antiretroviral therapy) regimen to Idvynso. A total of 551 participants were randomized (2:1) and were switched to once-daily Idvynso (n=366) or remained on their baseline ART (bART) (n=185). Randomization was stratified by bART. At baseline, participants had a mean age of 50 years (range: 18 to 83), 40% of participants were female, 39% were White, 45% were Black/African American, and 5% were Asian. A total of 15% identified as Hispanic/Latino. At enrollment, 64% of the participants were receiving integrase strand transfer inhibitor (INSTI)-based regimens, 5% protease inhibitor (PI)-based regimens (including combinations with INSTI), and 30% were receiving other regimens.
Monday, August 17, 2026
FDA Approves Revtorpyk (gedatolisib) for the Treatment of HR+/HER2-, PIK3CA Wild-Type Locally Advanced or Metastatic Breast Cancer
Celcuity Inc. (Nasdaq: CELC), a biotechnology company focused on developing and commercializing targeted therapies for multiple solid tumor indications, announced the U.S. Food and Drug Administration (“FDA”) approval of Revtorpyk (gedatolisib) for the treatment of patients with hormone receptor positive (“HR+”), human epidermal growth factor receptor 2 negative (“HER2-”), locally advanced or metastatic breast cancer without a PIK3CA mutation detected following progression on or after treatment with at least one line of endocrine therapy in the metastatic setting. Revtorpyk is the only inhibitor of class I PI3K isoforms (α, β, δ, γ) and mTOR complexes mTORC1 and mTORC2 to receive FDA approval.
- Revtorpyk is the first and only FDA-approved therapy that inhibits all class I PI3K isoforms (α, β, δ, γ) and mTOR complexes mTORC1 and mTORC2
- In the Phase 3 VIKTORIA-1 trial, Revtorpyk combined with palbociclib and fulvestrant and Revtorpyk combined with fulvestrant reduced the risk of disease progression or death by 76% and 67%, respectively, compared to fulvestrant among patients with PIK3CA wild-type advanced or metastatic breast cancer
“The PI3K/AKT/mTOR, or PAM, pathway is one of the most important targets in cancer, but comprehensively inhibiting it has stymied researchers and drug developers for nearly two decades,” said Brian Sullivan, CEO and co-founder of Celcuity. “Revtorpyk addressed this 20-year challenge by becoming the first pan-PI3K, mTORC1/2 inhibitor approved by the FDA. We are thankful for the opportunity to make this important new therapy available to patients with HR+/HER2- locally advanced or metastatic breast cancer.”
HR+/HER2- breast cancer is the most common subtype of breast cancer, accounting for approximately 70% of all breast cancers.1 Among this breast cancer subtype, approximately 60% have PIK3CA wild-type disease.2
“For patients with HR+/HER2- locally advanced or metastatic breast cancer, there is an urgent need for new treatment options that can meaningfully increase the likelihood of survival without disease progression or death,” said Sara Hurvitz, MD, Senior Vice President, Clinical Research Division, Fred Hutchinson Cancer Center, Smith Family Endowed Chair in Women’s Health and Professor and Head, Division of Hematology and Oncology, University of Washington, School of Medicine and co-principal investigator for the VIKTORIA-1 trial. “With the approval of Revtorpyk, oncologists now have an effective new treatment option for these patients.”
The approval of Revtorpyk is based on positive clinical results from the PIK3CA wild-type cohort of the Phase 3 VIKTORIA-1 trial, an open-label, global, randomized clinical trial evaluating the efficacy and safety of Revtorpyk plus fulvestrant, with or without palbociclib, for the treatment of patients with locally advanced or metastatic HR+/HER2- breast cancer following progression on or after CDK4/6 therapy and an aromatase inhibitor. In the VIKTORIA-1 trial, median progression free survival (“PFS”) with the Revtorpyk triplet (Revtorpyk plus palbociclib and fulvestrant) was 9.3 months versus 2.0 months with fulvestrant, an incremental improvement of 7.3 months (HR=0.24; 95% CI: 0.17-0.35; p<0.0001). The objective response rate (“ORR”) of the Revtorpyk triplet was 32% compared to 1% with fulvestrant and the median duration of response (“DOR”) was 17.5 months. For the Revtorpyk doublet (Revtorpyk plus fulvestrant), the median PFS was 7.4 months versus 2.0 months with fulvestrant, an incremental improvement of 5.4 months (HR=0.33; 95% CI: 0.24-0.48; p<0.0001). The ORR of the Revtorpyk doublet was 28% and the median DOR was 12.0 months. The median DOR was not determinable for fulvestrant because there was only one objective response.
“Today’s approval of Revtorpyk addresses a significant unmet need for the thousands of patients affected each year by HR+/HER2-, PIK3CA wild-type locally advanced or metastatic breast cancer whose disease has progressed after endocrine therapy,” said Igor Gorbatchevsky, MD, Chief Medical Officer of Celcuity. “We are deeply grateful to the patients and their caregivers, investigators and clinical study teams, and Celcuity team members who made this advancement possible.”
Celcuity anticipates commercial launch in late Q3 2026. Based on the company’s commitment to ensuring broad, affordable, and unrestricted patient access to Revtorpyk, we have designed a comprehensive patient support program. To make Revtorpyk available to patients prior to commercial launch, those who are eligible will be able to enroll in Celcuity’s expanded access program.
Celcuity plans to submit in Q3 2026 a supplemental New Drug Application (“sNDA”) to the FDA for Revtorpyk for the treatment of HR+/HER2-, PIK3CA mutated, locally advanced or metastatic breast cancer, following at least one line of endocrine therapy based on results from the mutant cohort of the Phase 3 VIKTORIA-1 trial. Results for this study were recently presented in a late-breaking abstract oral session at the 2026 American Society of Clinical Oncology (ASCO) Annual Meeting. Following the sNDA submission, Celcuity intends to submit VIKTORIA-1 data for marketing authorization of gedatolisib to other regulatory authorities around the world.
Revtorpyk is also being studied in the ongoing Phase 3 VIKTORIA-2 clinical trial incorporating two independent studies evaluating two separate patient cohorts with HR+/HER2- locally advanced or metastatic breast cancer who are treatment-naive in the advanced setting.
Revtorpyk is in development for the first-line treatment of HR+/HER2- locally advanced or metastatic breast cancer and for the second-line treatment of metastatic castration resistant prostate cancer.
Friday, July 17, 2026
FDA Approves Nereus (tradipitant) for the Prevention of Motion Sickness
Thursday, July 16, 2026
FDA Approves Zycubo (copper histidinate) for Children With Menkes Disease
Sentynl Therapeutics Inc. (“Sentynl”), a U.S.-based biopharmaceutical company wholly-owned by Zydus Lifesciences Limited (“Zydus Group”), announced that the U.S. Food and Drug Administration (“FDA”) has approved Zycubo® (copper histidinate) for the treatment of Menkes disease in pediatric patients. This is the first and only treatment approved for Menkes disease, a rare and fatal genetic condition, in the United States. Zycubo® is not indicated for the treatment of Occipital Horn Syndrome.
“Approval is a pivotal step towards achieving our goal of making a meaningful impact on patients, caregivers, and the rare disease community,” said Dr. Sharvil P. Patel, Managing Director, Zydus Lifesciences Limited. “This milestone marks a transformative moment for the Zydus Group and for families affected by Menkes disease. For the first time, patients have access to an approved therapy, offering hope where no options existed. We are proud to advance patient care and deliver innovative solutions to those who need them most.”
Menkes disease is a rare X-linked recessive pediatric disease caused by mutations of the copper transporter ATP7A encoded by the ATP7A gene. Patients with Menkes disease are born with the inability to absorb dietary copper and subsequently have impaired copper transport across the blood-brain barrier, and, until now, there has been no approved treatment in the United States. Zycubo® is a subcutaneous injectable formulation of copper histidinate that restores copper homeostasis and maintains copper levels in patients with Menkes disease.
“Menkes disease presents significant challenges for patients and their families. With no known cure, most untreated patients do not survive beyond three years of age,” said Matt Heck, CEO, Sentynl. “FDA’s approval serves as compelling affirmation that a safe and effective therapy is now available for patients living with this devastating disease.”
The approval is supported by positive topline clinical efficacy results for Zycubo, demonstrating statistically significant improvement in overall survival for Menkes disease subjects who received early treatment (“ET”) with Zycubo, compared to an untreated contemporaneous external control cohort, with a nearly 80% reduction in the risk of death. Median overall survival (“OS”) was 177.1 months for Zycubo ET cohort compared to 17.6 months for the untreated contemporaneous external control cohort.
The most common adverse reactions (incidence ≥7%) were pneumonia, viral infection, respiratory failure, seizure, bacterial infection, hemorrhage, hypotension, vomiting, tachycardia, pyrexia, volume depletion, fracture, dyspnea, transaminases elevation, diarrhea, fungal infection, anemia, and local administration reaction.
“This milestone represents the culmination of decades of research into better understanding and ultimately finding an effective treatment for Menkes disease,” said Dr. Stephen Kaler, a clinical genetics and genomics specialist at the Columbia University Medical Center. “Increased awareness of Menkes disease and rapid testing upon suspicion are critical, as beginning copper histidinate therapy in affected neonates has been shown to reduce symptoms and prolong life.”
Acquired from Cyprium Therapeutics in 2023, Sentynl has advanced Zycubo through the final stages of development with the FDA based on positive results from pivotal studies, receiving Breakthrough Therapy, Fast Track, Rare Pediatric Disease, and the FDA Orphan Drug Designations. Copper histidinate has also been granted Orphan Designation by the European Medicines Agency.
Wednesday, July 15, 2026
FDA Approves Zycubo (copper histidinate) for Children With Menkes Disease
Tuesday, July 14, 2026
FDA Approves Yuvezzi (carbachol and brimonidine tartrate) Ophthalmic Solution to Treat Presbyopia
Monday, July 6, 2026
FDA Approves Lynavoy (linerixibat) for Cholestatic Pruritus in Patients with Primary Biliary Cholangitis
Friday, May 29, 2026
FDA Approves Rhapsido (remibrutinib) for the Treatment of Chronic Spontaneous Urticaria
Wednesday, May 27, 2026
FDA Approves Foundayo (orforglipron), the Only GLP-1 Pill for Weight Loss That Can be Taken Any Time of Day Without Food or Water Restrictions
Wednesday, May 20, 2026
FDA Approves Jascayd (nerandomilast) to Treat Idiopathic Pulmonary Fibrosis
Thursday, May 14, 2026
FDA Approves Ferabright (ferumoxytol injection) Contrast Agent for Magnetic Resonance Imaging of the Brain
Tuesday, October 8, 2024
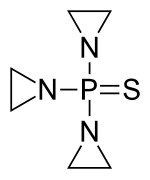
FDA Approves Tepylute (thiotepa) Ready-to-Dilute Injectable Formulation to Treat Breast Cancer and Ovarian Cancer
‘’This approval fulfills an unmet need by addressing the shortcomings and handling complexities of the current lyophilized powder formulation,” said Sharon Cunningham, Chief Executive Officer and Co-Founder of Shorla Oncology. “We have taken a vital oncology drug and made it easier for oncology clinics and hospitals to use, while also reducing medical personnel exposure to a hazardous drug.”
Tepylute, formerly SH-105, is the third FDA-approved drug for Shorla, and a significant milestone for the company as it seeks approval for several cancer-fighting drugs for the U.S. market.
“The approval of Tepylute represents an important milestone for Shorla as our first in-house developed NDA,” said Orlaith Ryan, Chief Technical Officer and Co-Founder of Shorla Oncology.
Tepylute is a liquid form of a well-established, standard of care oncology drug, thiotepa. The new formulation eliminates the need for complex and time-consuming reconstitution. It provides consistent dosing accuracy and allows for “just in time” preparation.2
“Among Tepylute’s many benefits, it removes the necessity to reconstitute which can introduce additional risks of drug preparation errors,” emphasized Rayna Herman, Chief Commercial Officer. “We look forward to providing an update on our launch plans for Tepylute in the near future.”
The American Cancer Society estimates that more than 300,000 women will be diagnosed with breast cancer in the U.S in 2024.3 About 19,680 women will be diagnosed with ovarian cancer in the United States.4
Shorla Oncology is currently marketing two products with a robust pipeline including SH-201, the first palatable oral liquid treatment for certain forms of leukemia and other cancers. In April, the company announced the FDA had accepted SH-201 for an NDA review with an expected action date of November 30, 2024.
‘’This approval fulfills an unmet need by addressing the shortcomings and handling complexities of the current lyophilized powder formulation,” said Sharon Cunningham, Chief Executive Officer and Co-Founder of Shorla Oncology. “We have taken a vital oncology drug and made it easier for oncology clinics and hospitals to use, while also reducing medical personnel exposure to a hazardous drug.”
Tepylute, formerly SH-105, is the third FDA-approved drug for Shorla, and a significant milestone for the company as it seeks approval for several cancer-fighting drugs for the U.S. market.
“The approval of Tepylute represents an important milestone for Shorla as our first in-house developed NDA,” said Orlaith Ryan, Chief Technical Officer and Co-Founder of Shorla Oncology.
Tepylute is a liquid form of a well-established, standard of care oncology drug, thiotepa. The new formulation eliminates the need for complex and time-consuming reconstitution. It provides consistent dosing accuracy and allows for “just in time” preparation.2
“Among Tepylute’s many benefits, it removes the necessity to reconstitute which can introduce additional risks of drug preparation errors,” emphasized Rayna Herman, Chief Commercial Officer. “We look forward to providing an update on our launch plans for Tepylute in the near future.”
The American Cancer Society estimates that more than 300,000 women will be diagnosed with breast cancer in the U.S in 2024.3 About 19,680 women will be diagnosed with ovarian cancer in the United States.4
Shorla Oncology is currently marketing two products with a robust pipeline including SH-201, the first palatable oral liquid treatment for certain forms of leukemia and other cancers. In April, the company announced the FDA had accepted SH-201 for an NDA review with an expected action date of November 30, 2024.